Key Findings
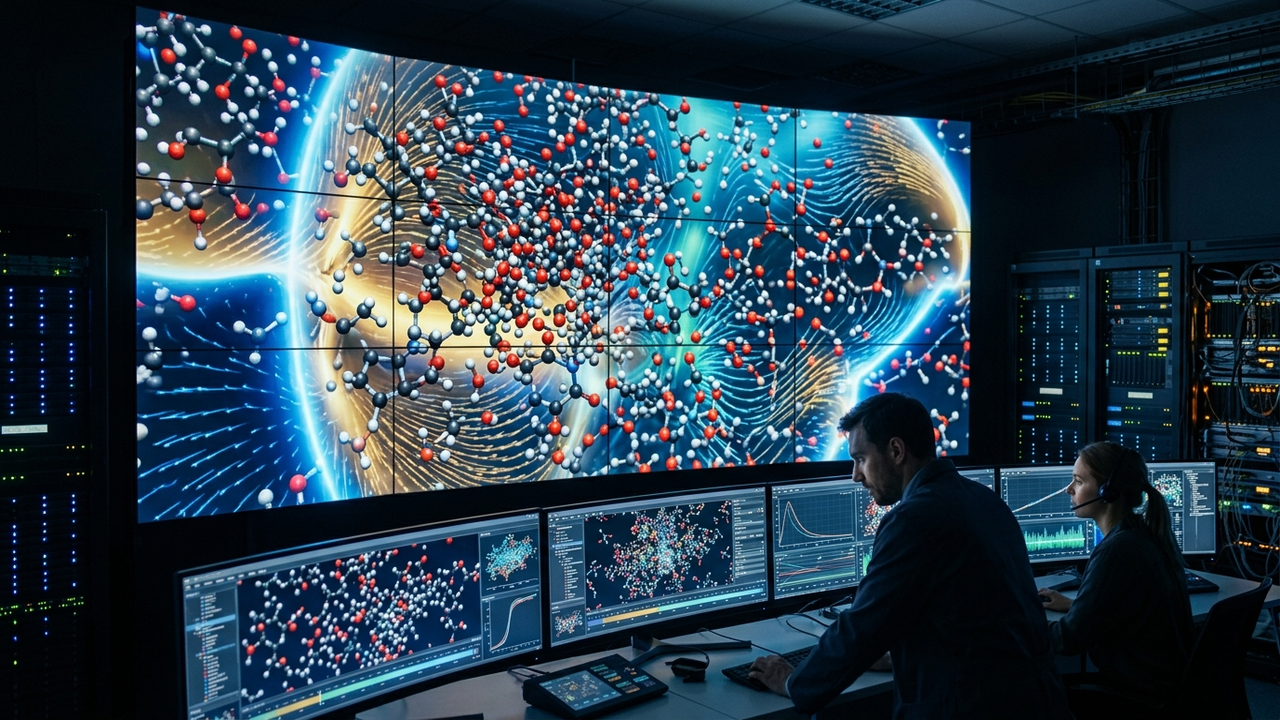
LAMMPS, a widely utilized open-source molecular dynamics (MD) simulator, has significantly enhanced its interoperability with advanced machine learning interatomic potentials (MLPs) such as DeePMD-kit and NequIP. This integration now allows researchers to directly incorporate high-fidelity MLPs, trained from quantum mechanical reference data, into LAMMPS simulations. This development achieves dramatically higher accuracy and computational efficiency compared to traditional MD simulations, representing a crucial advancement for understanding and designing complex material systems at the atomic level.
Technical / Clinical Details
Conventional MD simulations traditionally relied on classical force fields to model interatomic interactions, but their accuracy was limited by empirical parameterization and inherent difficulties in precisely capturing quantum mechanical phenomena. Conversely, quantum mechanical calculations (e.g., Density Functional Theory, DFT) offer high accuracy but are computationally expensive, precluding their application to large systems or long simulation times. MLPs like DeePMD-kit and NequIP are trained on small quantities of high-fidelity DFT data and can predict interatomic interactions with quantum-comparable accuracy. Their integration into LAMMPS enables MD simulations to maintain DFT-level accuracy while accelerating computation by several orders of magnitude. This capability allows for simulating phenomena previously computationally intractable, such as phase transitions, defect migration, catalytic reactions, and biomolecular dynamics, over more realistic time and spatial scales. Furthermore, repositories like OpenKIM offer curated libraries of various MLPs and their validation data, aiding in model selection and reliability assessment.
Background & Context
In numerous fields of materials science, chemistry, and biology, understanding dynamic atomic-level behavior is indispensable for developing new materials and designing functional molecules. Advancements in high-performance computing and machine learning technologies have dramatically expanded the accuracy and applicability of MD simulations. MLPs, in particular, are regarded as ‘game-changers’ as they overcome the limitations of classical force fields and extend quantum-level accuracy to larger systems. The enhanced interoperability of widely used platforms like LAMMPS with MLPs ensures that the broader research community can readily access these cutting-edge technologies. This accelerates material design and process optimization across academic research and industrial applications.
Strategic Significance & Outlook
The integration of LAMMPS with MLPs has the potential to usher in a new golden age for molecular dynamics simulations. Future developments are expected to focus on creating more universal MLPs capable of predicting material behavior in increasingly complex chemical systems and under extreme conditions. Moreover, synergy with AI-driven autonomous experimental systems will facilitate closed-loop material discovery platforms where simulation and experimentation are tightly coupled. This will dramatically boost the pace of discovery in materials science, contributing significantly to solving grand challenges in sustainable energy solutions, advanced medicine, and next-generation electronics.
Source: https://www.lammps.org/ecosystem/ml-interop/
Get our weekly technology intelligence — free
Receive an infographic that lets you judge at a glance whether each field’s analysis report is worth reading.
Subscribe Free — Weekly Tech Intelligence
By subscribing, you’ll receive Troy-Technical’s weekly technology intelligence newsletter.
- Your email and selected fields are used only to deliver the newsletter.
- We never share your information with third parties.
- You can unsubscribe anytime via the link in each email.
See our Privacy Policy for details.
Takes about a minute · Unsubscribe anytime

Comments