Key Findings
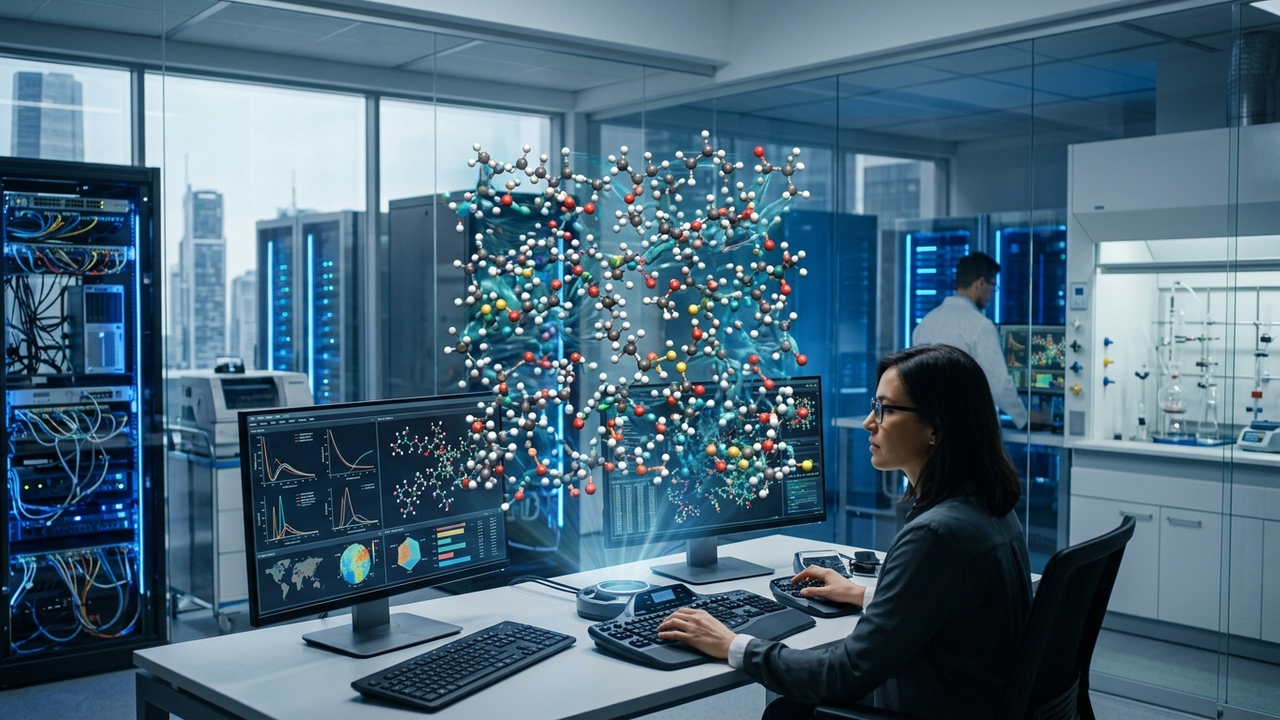
This research introduces GRACE-OFF (GRACE Organic Force Field), a high-performance machine-learned interatomic potential (MLIP) specifically designed for organic liquids, leveraging the Graph Atomic Cluster Expansion (GRACE) neural network architecture. GRACE-OFF enables molecular dynamics (MD) simulations to achieve accuracy comparable to quantum mechanical calculations while dramatically reducing computational costs. This breakthrough opens new avenues for detailed simulation of complex organic system dynamics over previously inaccessible time and spatial scales.
Technical / Clinical Details
MLIPs learn from quantum mechanical reference data to predict interatomic interaction energies and forces with high speed and precision. The GRACE architecture represents atomic local environments as graph structures, combining this with a many-body expansion (Atomic Cluster Expansion, ACE) framework to efficiently describe complex interatomic interactions. GRACE-OFF specifically adapts this GRACE architecture for predicting potential energy surfaces (PES) of organic molecules. Unlike traditional classical force fields, which often show reasonable accuracy only for limited molecular species or environments, GRACE-OFF reliably reproduces intermolecular interactions in organic liquids and accurately describes chemical reactions involving bond breaking and formation, even in systems with numerous constituent atoms and bonding types. This signifies a several-order-of-magnitude increase in computational speed while maintaining Density Functional Theory (DFT) accuracy, significantly broadening the applicability of molecular simulations in diverse fields such as materials design, drug discovery, and catalyst development.
Background & Context
Organic liquids are essential components in various technological and scientific applications, including battery electrolytes, solvents, fuels, and biological environments. However, accurately modeling their atomic-level behavior has been a long-standing challenge due to their diverse molecular structures and complex intermolecular interactions. Traditional simulation methods faced significant trade-offs between accuracy and computational cost, limiting large-scale and long-duration simulations. MLIPs, particularly models like GRACE-OFF which combine versatility and accuracy, offer a powerful solution to this challenge. They enable researchers and engineers to gain deeper insights for designing new organic materials, optimizing processes, and understanding biological phenomena. In industrial sectors, their adoption is expected to accelerate due to direct benefits in reducing new product development time and costs.
Strategic Significance & Outlook
The development of MLIPs for organic systems, such as GRACE-OFF, will further deepen the convergence of computational chemistry and materials science. Future research will likely focus on improving applicability across broader chemical spaces and extreme conditions (e.g., high temperature/pressure, supercritical fluids), as well as enhancing the ability to predict more complex reaction pathways. Furthermore, integration with autonomous experimental systems could lead to closed-loop material discovery processes where computationally predicted candidate materials are robotically synthesized and evaluated, with results fed back into the models. This is expected to be a driving force for generating innovative breakthroughs in fields such as drug discovery, polymer science, and organic electronics.
Get our weekly technology intelligence — free
Receive an infographic that lets you judge at a glance whether each field’s analysis report is worth reading.
Subscribe Free — Weekly Tech Intelligence
By subscribing, you’ll receive Troy-Technical’s weekly technology intelligence newsletter.
- Your email and selected fields are used only to deliver the newsletter.
- We never share your information with third parties.
- You can unsubscribe anytime via the link in each email.
See our Privacy Policy for details.
Takes about a minute · Unsubscribe anytime

Comments