TECHNOLOGY EXPLAINER
Off-Target Effects and Safety Assessment
— “not found” is a claim with a detection limit attached
At the heart of safety assessment for genome editing is the question of whether anything other than the intended site has been edited. In practice, though, the assessment is not a yes-or-no matter. In the public assessment report for CASGEVY, candidate sites were nominated by several methods and checked by deep sequencing, and no off-target editing was confirmedSourced. The prescribing information, on the other hand, says “the risk of unintended off-target editing due to genetic variants cannot be ruled out, and its clinical significance is unknown”Sourced. The two statements do not contradict each other. The sensitivity and the scope of what can be seen determine what the conclusion means.
- What off-target means, in three points
- There are four kinds of “unintended edit”
- A map of detection methods: nominating candidates and confirming them
- Our calculation: how far can “not found” be claimed?
- A materials engineer's view (1): the assessment system rests on reagents and controls
- A worked example: reading the CASGEVY assessment in the public assessment report
- Individual differences: genetic variants change the result
- How regulators think: FDA, PMDA and EMA
- Ways to reduce it: guides, enzymes, time and method
- A materials engineer's view (2): mitigation is a materials-design problem
- Open problems, and what is not yet settled
- Glossary / References / Claim-to-source audit
Sourced = stated in published material (link given)
Our calculation = a value this article derived from assumptions it states
Not yet confirmed = a plan or target with no confirmed track record
Structural readings and interpretations from an assessment-design standpoint are marked separately as Commentary.
This article is an explanation of assessment technology and materials technology. It does not judge the safety of any particular treatment, recommend a treatment, or rank products against one another, and it is not medical advice. Figures are quoted only where they appear in published material from regulators and in peer-reviewed papers, together with their conditions (cell type, time of assessment and so on). The mechanism of genome editing itself is covered in the companion article in this series, “Genome editing”.
1. What off-target means, in three points
Off-target means “an unintended edit that occurs at another site where a sequence similar to the intended one exists”.
- Why it happens: the PMDA Science Board report says that with CRISPR/Cas it is known that the guide RNA can bind even with up to 5 mismatches, which raises the likelihood of cleavage of unintended genes and the resulting insertions and deletionsSourced. A paper by Fu and colleagues also reported that mutations can be introduced at sites that differ from the target by up to 5 basesSourced
- What is of concern: the PMDA report sets out that the consequence of off-target activity of greatest concern is malignant transformation of cells, through activation of oncogenes or inactivation of tumour suppressor genes, and that because genome-editing changes are permanent the risk is greaterSourced
- How it is handled: FDA guidance recommends genome-wide analysis using several methods (in silico, biochemical and cell-based)Sourced. At approval, the FDA also made a 15-year observational study a mandatory post-marketing requirementSourced
“Not detected” means “not found at a frequency above the lower limit of sensitivity of that method”. In the EMA public assessment report, the thresholds for the confirmatory deep sequencing were 1% in the earlier study and 0.2% in the later ultra-deep studySourced. The PMDA report states that, because of the error rate of next-generation sequencers, it is extremely difficult to detect off-target activity occurring at a frequency of 0.1% or lowerSourced. This article uses the published figures to check what that lower limit means.
2. There are four kinds of “unintended edit”
The word “off-target” is in practice often used to lump together at least four different phenomena. FDA guidance lists as risks specific to genome editing off-target editing, unintended consequences of on-target editing, and unknown long-term effectsSourced. The four categories below are this article's own arrangement, based on published studies.
2. Large deletions and rearrangements at the target site
Kosicki and colleagues showed, in mouse embryonic stem cells, mouse haematopoietic progenitors and a human differentiated cell line, that double-strand breaks made by Cas9 are often repaired as deletions spanning several kb, and reported finding lesions distant from the break point and crossover events. The authors state that this genome damage, arising in dividing cells, could have pathogenic consequencesSourced. Cullot and colleagues reported, in cell lines and primary cells, that a single cut alone can cause megabase-scale chromosome truncation in a p53-dependent mannerSourced. The PMDA report also notes that large deletions spanning several kb, insertion of gene fragments and inversions have been reported during DSB repair, and explains that the cause is that the direction of repair is not determinedSourced.
3. Chromosome-level abnormalities
Leibowitz and colleagues showed, in model cells and in cells and loci relevant to clinical use, that Cas9 cutting can produce nuclear structural defects (micronuclei and chromatin bridges) and thereby chromothripsis, an extreme chromosomal rearrangement. The authors describe this as “an on-target consequence that has been overlooked until now” and state that in clinical applications the possibility of extensive chromosomal rearrangement should be considered and monitoredSourced. Zuccaro and colleagues reported that after Cas9 cutting in human embryos one or both arms of a chromosome can be lost (this is basic research on embryos, not a result for a therapeutic product)Sourced.
4. Selection by the cell's response
Haapaniemi and colleagues showed that editing with Cas9 triggers a p53-dependent DNA damage response and cell-cycle arrest, and that selection acts in favour of cells in which the p53 pathway is functional. The authors state that cell therapies should be developed with p53 function monitoredSourced. Ihry and colleagues obtained a mean indel efficiency above 80% in human pluripotent stem cells (hPSCs) by delivering Cas9 as an RNP, but showed that the DSBs made by Cas9 are toxic and kill many hPSCs, and that this response is P53/TP53-dependent. The authors say that, because hPSCs can acquire P53 mutations, cell-replacement therapy using Cas9-engineered hPSCs should proceed with caution and monitor P53 functionSourced. The PMDA report also mentions reports in which p53 mutations were detected in genome editing that used homology-directed repair, and says that the presence or absence of p53 and similar factors should be checkedSourced. This is a point of particular relevance to regenerative medicine using iPS cells (see this series' article on iPS cells).
3. A map of detection methods: nominating candidates and confirming them
Off-target assessment normally proceeds in two stages. First candidates are nominated for “where cutting could occur”, and then those candidates are read deeply and confirmed. The EMA public assessment report describes the CASGEVY assessment in exactly these two stages, “site nomination” and “site confirmation”Sourced. The PMDA report likewise sets out the structure: it is necessary to search for candidate sites across the whole human genome by experimental methods, not by in silico analysis alone, and then to check by WGS or amplicon sequencing whether cleavage or deletion actually occurs at those candidatesSourced.
| Method | Where it looks | What the original paper's abstract says |
|---|---|---|
| In silico (CCTop, CRISPOR, COSMID and others) | Computer (reference genome) | PMDA report: easy but inaccurate. EMA report: CCTop covers up to 5 mismatches, and COSMID can also search for candidates containing insertions and deletions (bulges) |
| Digenome-seq (Kim et al. 2015) | In vitro (purified genomic DNA) | Whole-genome sequencing of Cas9-digested genomes, looking for reads that share the same 5′ end at a cleavage site. Also validated sites with indels at below 0.1% |
| CIRCLE-seq (Tsai et al. 2017) | In vitro (circularised genomic DNA) | Highly sensitive; it can also identify off-targets arising from cell-type-specific single-nucleotide polymorphisms, allowing a specificity profile to be built for each individual |
| CHANGE-seq (Lazzarotto et al. 2020) | In vitro (tagmentation) | Scalable and automatable. In human primary T cells it identified 201,934 off-target sites across 110 targets at 13 loci. Off-targets in cells were 2 to 4 times more likely near active promoters, enhancers and transcribed regions. Human single-nucleotide variants had a significant effect on the activity of about 15.2% of sites |
| GUIDE-seq (Tsai et al. 2015) | In cells | Uses the integration of a double-stranded oligo DNA at cleavage sites. With 13 RGNs it found large variation in off-target activity. Many of the sites identified had not been detected by existing computational methods or ChIP-seq. Truncated guide RNAs greatly reduced off-targets |
| DISCOVER-seq (Wienert et al. 2019) | In cells and in vivo | Tracks recruitment of the DNA repair factor MRE11. Identified off-targets in cell lines, patient-derived iPS cells and in mouse tissue in vivo |
| Confirmation: deep sequencing | The actual edited cells | PMDA report: whole-genome sequencing is accurate but expensive and of low sensitivity. In methods that amplify candidate sites by PCR and read them deeply, sensitivity depends on read depth |
All Sourced (References 2, 4 and 8 to 12, including the assessment in Table 1 of the PMDA report). The order of the methods and the “Where it looks” grouping are this article's own arrangement.
4. Our calculation: how far can “not found” be claimed?
“Reading deeply” in the confirmation stage is a question of how many reads (sequenced fragments) are collected per candidate site. The EMA public assessment report sets the threshold for hybrid-capture deep sequencing at 1% (median coverage above 2,500) in the earlier study and 0.2% (ultra-deep, above 10,000×) in the later studySourced. From these numbers we calculate what a “threshold” means.
We make the following simplificationsOur calculation.
- Assumption 1: if a site has N reads and the true editing frequency at that site is f, the expected number of reads carrying the edit is N × f (assuming no sequencing errors)
- Assumption 2: when no read at all carrying the edit is found, the one-sided 95% upper bound on the true frequency is roughly 3 ÷ N (the “rule of three”)
- Assumption 3: the minimum recommended dose is 3×10⁶ CD34-positive cells/kg (prescribing information), with a body weight of 60 kg assumed. Cells are diploid and a single site is assessed
Calculation
- Expected reads: 1% × 2,500 = 25 reads / 0.2% × 10,000 = 20 reads / 0.1% × 10,000 = 10 reads / 0.05% × 10,000 = 5 reads
- Probability of reading 5 or more (Poisson distribution): about 97% if 10 are expected, about 56% if 5 are expected
- Upper bound when zero are found: 0 of 10,000 reads → 3 ÷ 10,000 = 0.03% (for 2,500 reads, about 0.12%)
- Converted to cell numbers: 3×10⁶ × 60 = 1.8×10⁸ cells → 3.6×10⁸ alleles. At a frequency of 0.2% that is about 7.2×10⁵ alleles; at 0.03%, about 1.1×10⁵ alleles
What can be read from this: a threshold of 0.2% at a depth of 10,000× corresponds to “20 reads expected”, which can be read as a design that secures enough reads to be told apart from errors (our interpretation). At the same time, it shows that “not detected” can be claimed only down to a frequency of about 0.03%, and even then several hundred thousand alleles' worth could remain hidden below detection.
Assumptions and limits: real sequencing has errors and, as the PMDA states, detection at 0.1% or lower is extremely difficultSourced. The calculation above is an idealisation that ignores errors, and is meant to show the order of magnitude of the upper bound. Whether a low-frequency site actually causes harm is a separate question that this calculation does not assess. The EMA report states that many of the candidates raised by GUIDE-seq were in introns or intergenic regionsSourced.
5. A materials engineer's view (1): the assessment system rests on reagents and controls
Off-target assessment is a “measurement”, but the measurement itself is a design problem of materials and reagents. The EMA public assessment report records a typical case.
- Concentration of the synthetic oligo used as a tag: in GUIDE-seq, a double-stranded oligo DNA (dsODN) is introduced into the cells at the same time, taken up at cleavage sites and then read out. According to the EMA report, in CD34-positive cells a dsODN concentration above 1 µM caused cytotoxicity and an increase in the number of cleavage sites detected. The first study (including concentrations above 1 µM) detected 857 sites, whereas a follow-up study in two other donors at 0.5 µM (keeping cell viability at 70% or more) raised 21 regions, with no overlap with the first studySourced. Because the donors and conditions differed, the cause cannot be pinned down, but the passage shows that the concentration of a reagent can change the content of the candidate list (our commentary)
- Controls tell you the background: in the same report's karyotype analysis, 60 of 240 metaphase cells were treated with SPY101-RNP and 5 had a chromosomal abnormality. Four of them were cells that had lost one or two chromosomes, and the same kind of abnormality was seen in control cells not electroporated; the applicant regards it as an artefact arising during sample preparation, with a frequency within the background rangeSourced. Without untreated or mock-treated controls, background noise would be misread as an effect of editing. FDA guidance also asks for appropriate controls to confirm the quality of the assay and the interpretability of the resultsSourced
- In vitro methods choose the “material” of the DNA: the PMDA report describes methods that use enzymatic reactions in a test tube as “highly sensitive and able to distinguish single-nucleotide polymorphisms” but “not reactions inside cells”Sourced. Purified genomic DNA has no chromosomal folding (chromatin), so such methods must be used with an understanding of what they reproduce and what they do not (our commentary)
- The amount of sample is itself a constraint: the FDA review summary says that only some genetic variants were actually tested at the candidate sites, giving sample limitations as the reasonSourced. Cells from patients are a finite material
Within the scope of this article we could not confirm whether public reference materials for off-target assessment (such as reference genome-edited samples) have been established. Each assessment is built on the premise that controls and acceptance criteria are set product by productNot yet confirmed.
6. A worked example: reading the CASGEVY assessment in the public assessment report
The EMA public assessment report (EPAR) describes in detail the off-target assessment of CASGEVY's guide RNA (SPY101). It is one of the few public documents that show how an actual assessment was put together and how far it looked, rather than the general picture (all figures are as recorded in the EPAR and are results of non-clinical studies using human haematopoietic stem and progenitor cells).
The same report also assesses large changes at the target site itself (on-target). Using long-range PCR and hybrid-capture sequence analysis, it records the mean rate of large insertions of more than 30 base pairs as 4.0% (LR-PCR) and 4.7% (hybrid capture), and the mean rate of large deletions of more than 30 base pairs as 6.3% (LR-PCR) and 4.9% (hybrid capture). Deletions reached up to about 7.5 kb and insertions up to 5.5 kbSourced. The significant difference in deletion rate between the two methods is explained by the fact that fragments containing large deletions tend to be amplified more efficiently by LR-PCRSourced. In other words, the choice of measurement method changes how the same phenomenon looks.
The FDA review summary states the non-clinical conclusion as follows: in CD34-positive cells from healthy donors, patients with SCD and patients with transfusion-dependent β-thalassaemia, candidate sites nominated by in silico and cell-based analyses were confirmed by hybrid capture, and no off-target editing was detected. It also states explicitly that in the analysis that took genetic variants into account, only candidate sites containing some of the variants were tested experimentally, owing to sample limitationsSourced.
7. Individual differences: genetic variants change the result
Candidates for off-target assessment are in most cases nominated against the reference genome (a standard sequence). But a patient's sequence is not identical to the reference genome.
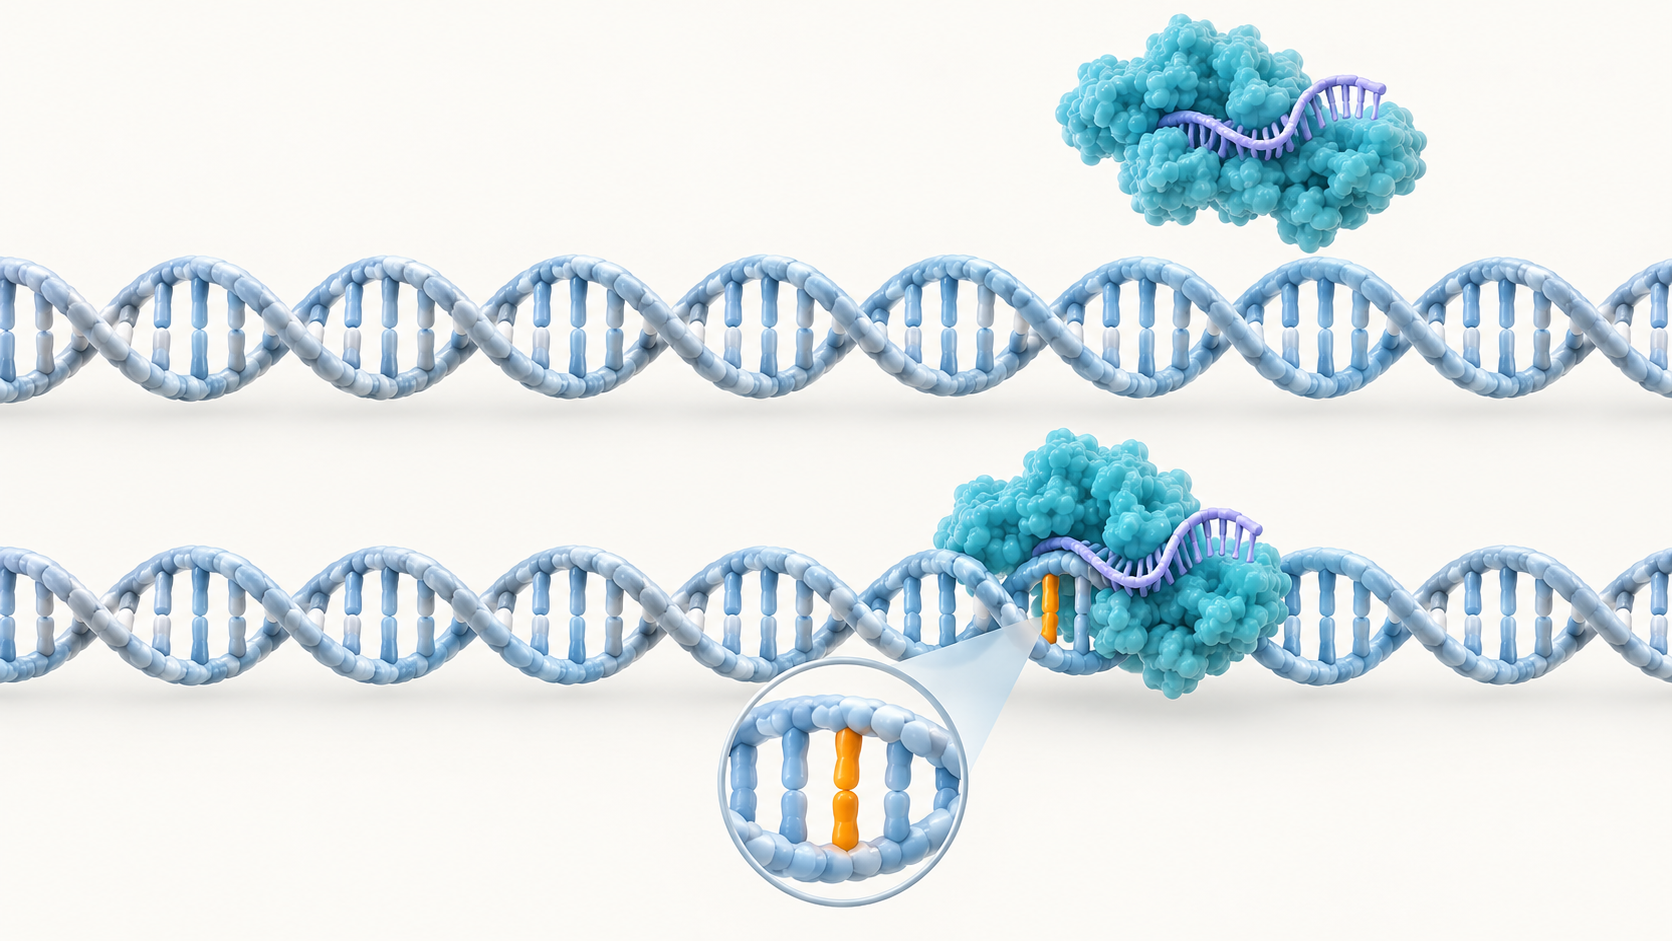 Top: reference genome (no PAM)
Bottom: individual variant (PAM can form)
Cancellieri and colleagues developed CRISPRme, a tool that searches for off-targets taking SNPs (single-nucleotide polymorphisms) and insertions and deletions into account, and tested it on a guide RNA targeting the BCL11A enhancer that had shown promise in clinical trials. They found that the strongest off-target candidate was created by an allele that is common in African-ancestry populations (minor allele frequency 4.5%), and that this allele newly generated a PAM sequenceSourced. The paper confirmed that SpCas9 produces strictly allele-specific indels and pericentromeric inversions in CD34-positive haematopoietic stem and progenitor cells, and reported that high-fidelity Cas9 mitigates this off-target. The authors state that “genetic variants should be considered as modifiers of the outcome of genome editing”, and expect that variant-aware off-target assessment will become indispensable in evaluating future therapeutic genome editingSourced.
The problem also appears in the regulators' documents.
- EMA report: it records that the homology-based candidate search was extended, using the 1000 Genomes Project, to SNPs with a minor allele frequency above 10% and later above 1%Sourced
- FDA review summary: it records that the analysis taking the genetic diversity of the patient population into account used a variant database built from the sequences of a limited number of people in the United States, and that some candidate sites created by variants were not tested experimentally. For this reason the FDA required a post-marketing bioinformatic study: specifically, repeating the in silico analysis with all variants that have an allele frequency of 0.5% or more in at least one of five continental populations (African, European, East Asian, South Asian and American), and confirmatory testing of any newly raised candidate sitesSourced
- Advisory committee: the FDA's Cellular, Tissue, and Gene Therapies Advisory Committee discussed the adequacy of the off-target safety assessment on 31 October 2023. The discussion touched on screening for a CPS1 variant and on additional analytical methods, and no voting question was putSourced. Whether this variant is the same as the allele reported by Cancellieri and colleagues cannot be determined from the published material
- Prescribing information: Section 5.4 states “the risk of unintended off-target editing in an individual's CD34-positive cells due to genetic variants cannot be ruled out; the clinical significance of off-target editing is unknown” (a major change in August 2025)Sourced
8. How regulators think: FDA, PMDA and EMA
Here we sort what has been covered so far by how each regulator's documents treat it. Note that guidance is a non-binding recommendation, and the requirements for each product are decided through individual consultation and review.
| Regulator / document | What it says about assessing off-target effects and chromosomal abnormalities |
|---|---|
| FDA Guidance (January 2024) | Carry out genome-wide analysis using several methods (in silico, biochemical and cell-based) to reduce bias. Where possible, analyse relevant human cells from several donors. Confirm with a method sensitive enough to detect low-frequency events, and for ex vivo edited products assess the final product from several donors. Include appropriate controls. Assess chromosomal abnormalities, insertions and deletions, tumourigenicity and insertional mutagenesis, and clonal expansion. Whether lot-release testing should include the off-target editing frequency or chromosomal rearrangements depends on the non-clinical results. Long-term follow-up of up to 15 years is recommended |
| FDA CASGEVY review summary (8 December 2023) | The most important identified risk is the possibility of off-target editing by CRISPR/Cas9, which is currently unknown. As conditions of approval it required an observational study following 250 SCD patients for 15 years (study completion planned for 31 December 2042, final report on 31 December 2043) and a bioinformatic study evaluating the effect of genetic diversity. As enhanced pharmacovigilance, expedited reporting within 15 days of secondary malignancy and clinical symptoms associated with off-target editing. A REMS (risk evaluation and mitigation strategy) was judged unnecessary |
| EMA CASGEVY EPAR (2024) | In non-clinical work, candidates were nominated by in silico analysis and GUIDE-seq and confirmed by hybrid capture. On-target effects and chromosomes were assessed by karyotyping, LR-PCR and similar methods. A conditional marketing authorisation with specific obligations, including a long-term follow-up study (up to 15 years) |
| PMDA Science Board report (February 2020) | Search for candidates across the whole human genome by experimental methods, not by in silico analysis alone, and confirm by whole-genome or amplicon sequencing. 0.1% or below is extremely difficult. Because of species differences, assessing off-target effects in animals is difficult and in vitro studies using human cells are needed. Chromosome analyses (G-banding, mFISH, CGH) each have limits to what they can detect. Haematopoietic stem cells and iPS / ES cells are assumed to carry a higher risk of malignant transformation than differentiated cells. It points to the need for long-term follow-up |
All Sourced (References 1 to 4). The dates of the post-marketing studies in the FDA review summary are plans as of December 2023, and current progress was not checked for this article (Not yet confirmed). The arrangement is this article's own and does not rank the regulators' requirements.
9. Ways to reduce it: guides, enzymes, time and method
Efforts to reduce off-target effects can be sorted into four levers (this article's arrangement). FDA guidance also asks that components such as the guide RNA be optimised to reduce off-target effectsSourced.
| Lever | What the paper abstract says |
|---|---|
| Truncated guide RNA (Fu et al. 2014) | Truncated guide RNAs whose target-complementary region is shorter than 20 bases can reduce undesired mutations at some off-target sites by 5,000-fold or more without compromising on-target editing efficiency. The GUIDE-seq report also found that truncated guide RNAs greatly reduced RGN-induced off-target DSBs |
| eSpCas9 (Slaymaker et al. 2016) | SpCas9 specificity improved by structure-guided protein engineering. In human cells it reduced off-targets while maintaining robust on-target cleavage |
| SpCas9-HF1 (Kleinstiver et al. 2016) | Designed to reduce non-specific DNA contacts. It matched wild-type on-target activity with more than 85% of the sgRNAs tested. With standard non-repetitive targets, almost all off-targets were undetectable by genome-wide break-capture methods and targeted sequencing. With repetitive targets, most of those seen with wild type were not detected |
| HiFi Cas9 (Vakulskas et al. 2018) | Existing high-fidelity variants lost on-target activity in therapeutically relevant RNP delivery. A bacterial screen yielded the point mutation p.R691A, which reduced off-target editing while keeping on-target activity. Gene targeting was confirmed at five loci in human CD34-positive cells and T cells |
| RNP delivery (Kim et al. 2014) | Because RNP cleaves almost immediately after delivery and is rapidly degraded in cells, it reduces off-target mutations compared with plasmid delivery. It is also gentler on pluripotent stem cells |
| Prime editing (Anzalone et al. 2019) | At known Cas9 off-target sites it showed much lower off-target editing than Cas9 nuclease |
| Concerns specific to base editing | Zuo et al.: using the GOTI method, off-target SNVs from Cas9 and adenine base editing were close to the spontaneous mutation rate, but cytosine base editing introduced SNVs at more than 20 times the frequency (mouse embryos). Grünewald et al.: a CBE using rat APOBEC1 caused RNA C-to-U editing at tens of thousands of sites, in 38 to 58% of expressed genes, which engineered variants cut by more than 390-fold and more than 3,800-fold. Adenine base editing can also cause RNA editing |
All Sourced (abstracts of References 7, 8 and 20 to 28). Figures are results under each paper's experimental conditions (cultured cells, mouse embryos and so on).
10. A materials engineer's view (2): mitigation is a materials-design problem
The paper by Vakulskas and colleagues holds a lesson familiar to anyone who develops materials. Existing high-fidelity Cas9 variants lost on-target activity in the form of delivery relevant to therapy, the RNPSourced. So they built a bacterial screening system to look for variants that keep their activity as an RNP, and obtained a single point mutation (R691A)Sourced. In other words, the performance of an enzyme has to be evaluated in the form in which it will be used as a product (RNP), not by its value when expressed from a plasmid (our commentary).
- A “fewer contacts” design: SpCas9-HF1 is designed to reduce non-specific DNA contactsSourced. It can be read as a design that strips away binding margin so that cleavage happens only with a perfect match to the target (our interpretation). In the language of materials, it is a design that sharpens the threshold
- Sequence length as a parameter: shortening the target-complementary region of the guide RNA from 20 bases is an operation on a design parameter of a chemically synthesised oligonucleotide. According to the EMA report, CASGEVY's guide RNA is a 100-mer synthetic product whose purity was raised by optimising synthesis and purification conditionsSourced. Both sequence design and impurity control bear on the specificity of the product (our commentary)
- RNP as a “single-use form”: the advantage of RNP is that it is degraded soon after delivery, but this also limits the cell types and conditions in which it can be delivered. Electroporation conditions, the order of mixing with the cells, and the time between preparing the RNP and using it are treated as process parameters in the EMA report tooSourced (the RNP is prepared immediately before electroporation)
Whether the Cas9 used in CASGEVY is a high-fidelity type could not be confirmed in the regulatory documents this article consulted (the FDA review summary, the EMA report and the prescribing information). What the regulatory documents state goes only as far as Cas9 derived from Streptococcus pyogenesNot yet confirmed.
11. Open problems, and what is not yet settled
(1) A detection limit is also a limit of the method
As Section 4 showed, frequencies below the threshold are, in effect, “not seen”. The PMDA says detection at 0.1% or lower is extremely difficult. The more sensitivity is raised, the more samples (cell numbers and DNA amounts) and money are neededSourced.
(2) How far individual differences can be covered
The FDA review summary raised the problem that the variant database rests on a limited number of people and asked for additional studies. Which populations and what frequency of variants should be included in the assessment is an area that regulators are still discussing (the FDA gave an allele frequency of 0.5% or more as an example)Sourced.
(3) Standardising the assessment methods
FDA guidance recommends using several methods together rather than a single one and does not name a particular “standard method”. In the EMA report too, the confirmation threshold changed from 1% to 0.2% between studiesSourced. Whether common reference materials and standard procedures for off-target assessment exist could not be confirmed within the scope of this article.
(4) Assessing in vivo editing
The PMDA report says that, because the base sequences of the genome differ between humans and animals, it is difficult to assess off-target activity in animals, and detailed analysis by in vitro studies using human cells is needed. For in vivo editing, analysis using primary cells is preferable to established cell lines, and cells derived from iPS or ES cells are also thought to be usefulSourced. DISCOVER-seq has been shown to identify sites in the mouse body (editing by adenovirus), but methods for detection in the human body are still being establishedSourced.
(5) Long-term effects
The prescribing information says the clinical significance of off-target editing is unknown. FDA guidance recommends follow-up of up to 15 years, and a 15-year long-term follow-up study for CASGEVY is registered (ClinicalTrials.gov NCT04208529)Sourced. Results from the post-marketing studies will come in several years to over ten years at the earliestNot yet confirmed.
- There are four kinds of “unintended edit”. Off-target editing at similar sequences, large deletions and rearrangements at the target, chromosome-level abnormalities, and selection through the cell response (p53)Sourced
- Assessment has two stages. Candidates are nominated (in silico, in vitro, in cells) and then confirmed by deep sequencing. Each method sees a different rangeSourced
- “Not found” is a claim with a detection limit attached. A threshold of 0.2% at a depth of 10,000× means 20 reads expected, and even zero found gives an upper bound of about 0.03%, leaving several hundred thousand alleles' worth hidden below itOur calculation
- In the non-clinical assessment of CASGEVY, no off-target editing was confirmed. The prescribing information nonetheless says the possibility of off-target editing due to genetic variants cannot be ruled outSourced
- Individual differences change the result. A case in which a variant creates a PAM has been reported, and the FDA required a bioinformatic studySourced
- Mitigation is a materials-design problem. There are levers in the guide, the enzyme, the time and the method, and the performance of an enzyme has to be assessed in the form in which it will be used (RNP)Sourced
- Long-term follow-up is a precondition. The FDA recommends follow-up of up to 15 years and also required a 15-year observational study at approvalSourced
12. Glossary
- Off-target
- Unintended editing at a similar sequence other than the intended site.
- On-target
- The intended site. Large deletions or rearrangements there also count as “unintended consequences”.
- Nomination
- The stage of listing candidate off-target sites.
- In silico
- Analysis on a computer. Candidates are predicted from sequence similarity in the reference genome.
- GUIDE-seq
- A method that has double-stranded oligo DNA taken up at cleavage sites so that the sites cut inside cells can be read out.
- CIRCLE-seq / CHANGE-seq
- Methods that cut circularised or tagmented genomic DNA with Cas9 in a test tube and read the cleavage sites.
- Digenome-seq
- A method that sequences the whole genome after Cas9 digestion to find cleavage sites.
- DISCOVER-seq
- A method that tracks recruitment of the DNA repair factor MRE11 to find cleavage sites in cells and in vivo.
- Hybrid capture
- A method that enriches only the regions of interest with probes matching the candidate sites and reads them deeply.
- Coverage (depth)
- How many reads cover a given position. The deeper, the easier it is to find low-frequency changes.
- Chromothripsis
- Extensive rearrangement confined to one or a few chromosomes.
- p53
- A tumour suppressor gene product that triggers cell-cycle arrest or cell death in response to DNA damage.
- Minor allele frequency (MAF)
- The frequency, in a population, of the less common allele.
- High-fidelity Cas9
- Cas9 engineered to reduce off-target effects (eSpCas9, SpCas9-HF1, HiFi Cas9 and others).
- Karyotyping
- A method that observes metaphase chromosomes to check for abnormalities in number and large structural changes.
- Long-term follow-up
- Following safety over a long period after administration. For genome editing the FDA recommends up to 15 years.
13. References (primary sources)
- US Food and Drug Administration (FDA) “Human Gene Therapy Products Incorporating Human Genome Editing — Guidance for Industry”, January 2024 (PDF) — fda.gov
- PMDA Science Board “Report on considerations for the quality and safety of gene therapy products using genome editing technology” (in Japanese), 7 February 2020 (PDF; the table of contents is at the start of the file) — pmda.go.jp
- US FDA CBER “Summary Basis for Regulatory Action — CASGEVY (BLA STN 125787)”, 8 December 2023 (PDF) — fda.gov
- EMA “Casgevy — EPAR Public assessment report”, EMA/6332/2024 (PDF) — ema.europa.eu
- EMA “Casgevy — European public assessment report (EPAR)” product page — ema.europa.eu
- US FDA “CASGEVY Prescribing Information”, revised July 2026 (PDF) — fda.gov
- Fu Y et al. “Improving CRISPR-Cas nuclease specificity using truncated guide RNAs”, Nat Biotechnol 32(3):279-284 (2014). doi:10.1038/nbt.2808 — pubmed.ncbi.nlm.nih.gov
- Tsai SQ et al. “GUIDE-seq enables genome-wide profiling of off-target cleavage by CRISPR-Cas nucleases”, Nat Biotechnol 33(2):187-197 (2015). doi:10.1038/nbt.3117 — pubmed.ncbi.nlm.nih.gov
- Tsai SQ et al. “CIRCLE-seq: a highly sensitive in vitro screen for genome-wide CRISPR-Cas9 nuclease off-targets”, Nat Methods 14(6):607-614 (2017). doi:10.1038/nmeth.4278 — pubmed.ncbi.nlm.nih.gov
- Lazzarotto CR et al. “CHANGE-seq reveals genetic and epigenetic effects on CRISPR-Cas9 genome-wide activity”, Nat Biotechnol 38(11):1317-1327 (2020). doi:10.1038/s41587-020-0555-7 — pubmed.ncbi.nlm.nih.gov
- Kim D et al. “Digenome-seq: genome-wide profiling of CRISPR-Cas9 off-target effects in human cells”, Nat Methods 12(3):237-243 (2015). doi:10.1038/nmeth.3284 — pubmed.ncbi.nlm.nih.gov
- Wienert B et al. “Unbiased detection of CRISPR off-targets in vivo using DISCOVER-Seq”, Science 364(6437):286-289 (2019). doi:10.1126/science.aav9023 — pubmed.ncbi.nlm.nih.gov
- Kosicki M et al. “Repair of double-strand breaks induced by CRISPR-Cas9 leads to large deletions and complex rearrangements”, Nat Biotechnol 36(8):765-771 (2018). doi:10.1038/nbt.4192 — pubmed.ncbi.nlm.nih.gov
- Cullot G et al. “CRISPR-Cas9 genome editing induces megabase-scale chromosomal truncations”, Nat Commun 10:1136 (2019). doi:10.1038/s41467-019-09006-2 — pubmed.ncbi.nlm.nih.gov
- Leibowitz ML et al. “Chromothripsis as an on-target consequence of CRISPR-Cas9 genome editing”, Nat Genet 53(6):895-905 (2021). doi:10.1038/s41588-021-00838-7 — pubmed.ncbi.nlm.nih.gov
- Zuccaro MV et al. “Allele-specific chromosome removal after Cas9 cleavage in human embryos”, Cell 183(6):1650-1664 (2020). doi:10.1016/j.cell.2020.10.025 — pubmed.ncbi.nlm.nih.gov
- Haapaniemi E et al. “CRISPR-Cas9 genome editing induces a p53-mediated DNA damage response”, Nat Med 24(7):927-930 (2018). doi:10.1038/s41591-018-0049-z — pubmed.ncbi.nlm.nih.gov
- Ihry RJ et al. “p53 inhibits CRISPR-Cas9 engineering in human pluripotent stem cells”, Nat Med 24(7):939-946 (2018). doi:10.1038/s41591-018-0050-6 — pubmed.ncbi.nlm.nih.gov
- Cancellieri S et al. “Human genetic diversity alters off-target outcomes of therapeutic gene editing”, Nat Genet 55(1):34-43 (2023). doi:10.1038/s41588-022-01257-y — pubmed.ncbi.nlm.nih.gov
- Slaymaker IM et al. “Rationally engineered Cas9 nucleases with improved specificity”, Science 351(6268):84-88 (2016). doi:10.1126/science.aad5227 — pubmed.ncbi.nlm.nih.gov
- Kleinstiver BP et al. “High-fidelity CRISPR-Cas9 nucleases with no detectable genome-wide off-target effects”, Nature 529(7587):490-495 (2016). doi:10.1038/nature16526 — pubmed.ncbi.nlm.nih.gov
- Vakulskas CA et al. “A high-fidelity Cas9 mutant delivered as a ribonucleoprotein complex enables efficient gene editing in human hematopoietic stem and progenitor cells”, Nat Med 24(8):1216-1224 (2018). doi:10.1038/s41591-018-0137-0 — pubmed.ncbi.nlm.nih.gov
- Kim S et al. “Highly efficient RNA-guided genome editing in human cells via delivery of purified Cas9 ribonucleoproteins”, Genome Res 24(6):1012-1019 (2014). doi:10.1101/gr.171322.113 — pubmed.ncbi.nlm.nih.gov
- Komor AC et al. “Programmable editing of a target base in genomic DNA without double-stranded DNA cleavage”, Nature 533(7603):420-424 (2016). doi:10.1038/nature17946 — pubmed.ncbi.nlm.nih.gov
- Gaudelli NM et al. “Programmable base editing of A•T to G•C in genomic DNA without DNA cleavage”, Nature 551(7681):464-471 (2017). doi:10.1038/nature24644 — pubmed.ncbi.nlm.nih.gov
- Zuo E et al. “Cytosine base editor generates substantial off-target single-nucleotide variants in mouse embryos”, Science 364(6437):289-292 (2019). doi:10.1126/science.aav9973 — pubmed.ncbi.nlm.nih.gov
- Grünewald J et al. “Transcriptome-wide off-target RNA editing induced by CRISPR-guided DNA base editors”, Nature 569(7756):433-437 (2019). doi:10.1038/s41586-019-1161-z — pubmed.ncbi.nlm.nih.gov
- Anzalone AV et al. “Search-and-replace genome editing without double-strand breaks or donor DNA”, Nature 576(7785):149-157 (2019). doi:10.1038/s41586-019-1711-4 — pubmed.ncbi.nlm.nih.gov
- ClinicalTrials.gov “A Long-term Follow-up Study in Participants Who Received CTX001 (NCT04208529)” — clinicaltrials.gov
14. Claim-to-source audit
| Claim in the text | Basis | Label |
|---|---|---|
| What the FDA guidance says: the risks specific to genome editing (off-target editing, unintended consequences of on-target editing, unknown long-term effects); that a DSB can cause chromosomal rearrangements (including translocations); genome-wide analysis by several methods (in silico, biochemical, cell-based); obtaining relevant human cells from several donors; verifying with a method sensitive enough to detect low-frequency events; assessing the final product from several donors for ex vivo edited products; including appropriate controls; assessing chromosomal abnormalities, insertions and deletions, tumourigenicity and clonal expansion; the possibility of including off-target editing frequency or chromosomal rearrangement in lot-release testing; optimisation of components; long-term follow-up of up to 15 years; minimising duration | FDA guidance (January 2024). Reference 1 https://www.fda.gov/media/156894/download | Sourced |
| What the PMDA report says: that the guide RNA can bind with up to 5 mismatches; that the consequence of concern from off-target activity is malignant transformation; that large deletions and the like that conventional assessment cannot detect have been reported with DSBs; searching the whole human genome by experimental methods in addition to in silico; confirmation by whole-genome or amplicon sequencing; that 0.1% or below is extremely difficult because of the NGS error rate; the table of detection methods (in silico is easy but inaccurate, whole-genome is accurate but expensive and of low sensitivity, in vitro is highly sensitive and distinguishes SNPs but is not in cells); that assessment in animals is difficult; large deletions, insertions and inversions of several kb; the limits of G-banding, mFISH and CGH; reports of p53 mutations; that haematopoietic stem cells and iPS / ES cells are assumed to carry higher risk of malignant transformation; use of primary cells or iPS / ES-derived cells for in vivo editing; long-term follow-up | PMDA Science Board report (7 February 2020; in Japanese). Reference 2 https://www.pmda.go.jp/files/000265952.pdf | Sourced |
| What the FDA review summary says: that the main risk is the possibility of off-target editing by CRISPR/Cas9, currently unknown; that in non-clinical work candidate sites were confirmed by hybrid capture and no off-target editing was detected; that the analysis reflecting genetic diversity relied on a database of a limited number of people and only some variants were tested experimentally (sample limitations); the post-marketing bioinformatic study (analysis with variants at 0.5% or more allele frequency in at least one of five continental populations, and confirmatory testing); the observational study following 250 SCD patients for 15 years (completion planned 31 December 2042, final report 31 December 2043); reporting within 15 days of secondary malignancy and clinical symptoms associated with off-target editing; no REMS; the advisory committee (31 October 2023, no voting question, discussion including screening for a CPS1 variant) | FDA review summary (SBRA, 8 December 2023). Reference 3 https://www.fda.gov/media/175179/download?attachment | Sourced |
| What the EPAR says: the two stages of nomination and confirmation; the in silico tools (CCTop, CRISPOR, COSMID) and mismatch numbers; the GUIDE-seq conditions (toxicity above 1 µM dsODN, 857 sites, viability of 70% or more at 0.5 µM, 21 regions in two other donors, no overlap, 5,007 sites in silico, 52 regions to 142 candidates); 64 regions in patient cells; hybrid capture (threshold 1% with median coverage above 2,500, later 0.2% at over 10,000×, 223 sites, 4 donors); 4 regions in patient cells, 2 nominally significant, and the applicant's reasons for not regarding them as true off-targets; assessment of 7 lots; no off-target editing confirmed in any study; the rates of large insertions and deletions (LR-PCR 4.0% and 6.3%, hybrid capture 4.7% and 4.9%, up to about 7.5 kb and 5.5 kb); karyotyping (60 of 240 cells treated, 5 abnormal, 4 with chromosome loss also present in controls); SNPs above 10% and above 1% MAF; that many candidates were in introns or intergenic regions; the conditional authorisation and the long-term follow-up obligation; that the guide RNA is a 100-mer whose purity was raised by optimising synthesis and purification conditions; that the RNP is prepared immediately before electroporation | EMA public assessment report EMA/6332/2024. Reference 4 https://www.ema.europa.eu/en/documents/assessment-report/casgevy-epar-public-assessment-report_en.pdf | Sourced |
| Section 5.4 of the CASGEVY prescribing information (the risk of off-target editing due to genetic variants cannot be ruled out and its clinical significance is unknown; a major change in August 2025), and the minimum recommended dose of 3×10⁶ CD34-positive cells/kg | FDA prescribing information. Reference 6 https://www.fda.gov/media/174615/download?attachment | Sourced |
| That mutations can be introduced at sites that differ from the target by up to 5 bases, and that truncated guide RNAs can reduce some off-target mutations by 5,000-fold or more | Abstract of Fu et al. (2014). Reference 7 https://pubmed.ncbi.nlm.nih.gov/24463574/ | Sourced |
| The principles and results of GUIDE-seq, CIRCLE-seq, CHANGE-seq, Digenome-seq and DISCOVER-seq (as described in the table) | Tsai et al. (2015). Reference 8 https://pubmed.ncbi.nlm.nih.gov/25513782/; Tsai et al. (2017). Reference 9 https://pubmed.ncbi.nlm.nih.gov/28459458/; Lazzarotto et al. (2020). Reference 10 https://pubmed.ncbi.nlm.nih.gov/32541958/; Kim et al. (2015). Reference 11 https://pubmed.ncbi.nlm.nih.gov/25664545/; Wienert et al. (2019). Reference 12 https://pubmed.ncbi.nlm.nih.gov/31000663/ | Sourced |
| Deletions of several kb and complex rearrangements at the target site, megabase-scale chromosome truncation (p53-dependent), chromothripsis, and loss of a chromosome arm in human embryos | Kosicki et al. (2018). Reference 13 https://pubmed.ncbi.nlm.nih.gov/30010673/; Cullot et al. (2019). Reference 14 https://pubmed.ncbi.nlm.nih.gov/30850590/; Leibowitz et al. (2021). Reference 15 https://pubmed.ncbi.nlm.nih.gov/33846636/; Zuccaro et al. (2020). Reference 16 https://pubmed.ncbi.nlm.nih.gov/33125898/ | Sourced |
| The p53-dependent DNA damage response and selection caused by Cas9 editing; the toxicity of Cas9 DSBs in pluripotent stem cells and their P53 dependence; a mean indel efficiency above 80%; the recommendation to monitor p53 function | Haapaniemi et al. (2018). Reference 17 https://pubmed.ncbi.nlm.nih.gov/29892067/; Ihry et al. (2018). Reference 18 https://pubmed.ncbi.nlm.nih.gov/29892062/ | Sourced |
| That the strongest off-target candidate for a BCL11A-enhancer-targeting guide RNA arises from a PAM created by an allele common in African-ancestry populations (MAF 4.5%); allele-specific indels and pericentromeric inversions; mitigation by high-fidelity Cas9 | Abstract of Cancellieri et al. (2023). Reference 19 https://pubmed.ncbi.nlm.nih.gov/36522432/ | Sourced |
| The descriptions and figures for eSpCas9, SpCas9-HF1, HiFi Cas9 (R691A) and RNP delivery | Slaymaker et al. (2016). Reference 20 https://pubmed.ncbi.nlm.nih.gov/26628643/; Kleinstiver et al. (2016). Reference 21 https://pubmed.ncbi.nlm.nih.gov/26735016/; Vakulskas et al. (2018). Reference 22 https://pubmed.ncbi.nlm.nih.gov/30082871/; Kim et al. (2014). Reference 23 https://pubmed.ncbi.nlm.nih.gov/24696461/ | Sourced |
| Off-target effects specific to base editing and prime editing (CBE SNVs more than 20-fold in mouse embryos, RNA editing, reductions of more than 390-fold and 3,800-fold, and fewer off-targets with prime editing) | Zuo et al. (2019). Reference 26 https://pubmed.ncbi.nlm.nih.gov/30819928/; Grünewald et al. (2019). Reference 27 https://pubmed.ncbi.nlm.nih.gov/30995674/; Anzalone et al. (2019). Reference 28 https://pubmed.ncbi.nlm.nih.gov/31634902/; Komor et al. (2016). Reference 24 https://pubmed.ncbi.nlm.nih.gov/27096365/; Gaudelli et al. (2017). Reference 25 https://pubmed.ncbi.nlm.nih.gov/29160308/ | Sourced |
| That a 15-year long-term follow-up study of CASGEVY is registered on ClinicalTrials.gov | ClinicalTrials.gov NCT04208529. Reference 29 https://clinicaltrials.gov/study/NCT04208529 | Sourced |
| The planned completion dates of the post-marketing studies in the FDA review summary, and their results (which will come several years to over ten years from now) | Plans as of December 2023; current progress was not checked by this article. No results are available | Not yet confirmed |
| Whether the Cas9 used in CASGEVY is a high-fidelity type; whether public reference materials for off-target assessment exist; whether common standard procedures exist | Not stated in this article because they could not be confirmed in the regulatory documents and literature consulted | Not yet confirmed |
| Expected reads (25, 20, 10 and 5), Poisson probabilities of reading 5 or more (about 97% and about 56%), rule-of-three upper bounds (about 0.03% and about 0.12%), the conversions to 1.8×10⁸ cells, 3.6×10⁸ alleles, and 7.2×10⁵ and 1.1×10⁵ alleles | Our calculation. An idealisation that ignores sequencing errors and assumes a 60 kg body weight, diploid cells and a single assessed site; it does not show actual detection ability or safety | Our calculation |
| Sorting “unintended edits” into four kinds. Sorting mitigation into four levers. The relation between dsODN concentration and the candidate list, the role of controls, the “sharpened threshold” design of high-fidelity enzymes, and the reading that assessment in the RNP form is needed. The interpretation that 20 reads expected is enough to be told apart from errors | This article's own framing and commentary based on published content. Not views expressed by the institutions | Commentary |
| That Figs. 1, 2, 3, 4, 6 and 7 are explanatory drawings, and that the hero image and Fig. 5 are AI-generated images | A note by this article | Commentary |
Last updated 1 October 2026. Sources are limited to primary material (published documents from regulators, public assessment reports and peer-reviewed papers). Because the article includes structural readings and interpretations from an assessment-design standpoint, those are marked as Commentary and kept separate from sourced fact. Whether the Cas9 in CASGEVY is a high-fidelity type, whether public reference materials or common standard procedures for off-target assessment exist, and the results of the post-marketing studies are not asserted, because they could not be confirmed in published primary sources. The calculation in Section 4 is an idealisation that ignores errors and does not guarantee real detection ability. All figures are concept diagrams for explanation. Figs. 1, 2, 3, 4, 6 and 7 are vector drawings; the hero image and Fig. 5 are AI-generated images, and none of them shows a real molecular structure, a microscope image, sequence data or an actual product.