Key Findings
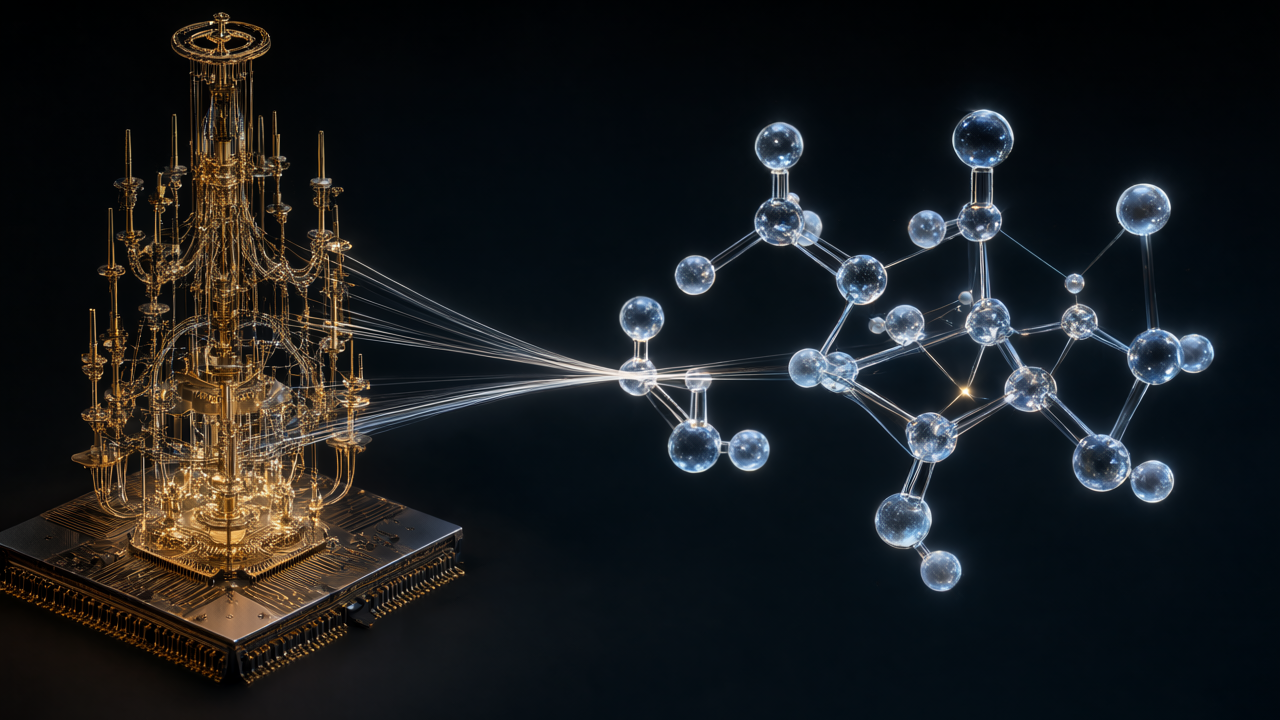
A novel framework has been proposed that effectively integrates sparse, high-fidelity data from quantum computing-based electronic structure calculations with universal machine-learning interatomic potentials (uMLIPs). This method allows for the fine-tuning of pre-trained density functional theory (DFT)-based uMLIPs using quantum-computed reference energies, dramatically enhancing the accuracy of atomistic simulations for complex phenomena like chemical reactions and adsorption.
Technical / Clinical Details
The core of this framework involves using a small but highly accurate dataset obtained from quantum computing to refine a uMLIP that has been pre-trained on a broader, less precise DFT dataset. Quantum computing, while offering unparalleled accuracy in electronic structure calculations, is computationally expensive and currently limited to smaller systems. By judiciously selecting a few critical data points (e.g., transition states, adsorption energies) for quantum calculation and using them to fine-tune the uMLIP, the model gains high-precision insights without incurring prohibitive computational costs for large-scale simulations. This hybrid approach allows for the simulation of complex atomic processes, such as catalytic reactions on surfaces or dynamic phase transitions, with an accuracy level previously unattainable by either method alone for large systems.
Background & Context
The quest for accurate and efficient atomistic simulations is central to materials science and engineering. While DFT provides a good balance of accuracy and computational cost, its limitations become apparent in highly correlated systems or when simulating large systems for extended periods. Machine learning interatomic potentials (MLIPs) offer computational speed but often lack the quantum-level accuracy for subtle electronic effects. Quantum computing, despite its promise, is not yet mature enough for routine large-scale materials simulations. This research elegantly combines the strengths of quantum computing (accuracy for key points) with MLIPs (computational efficiency for large systems) to overcome these individual limitations, addressing a long-standing challenge in computational materials science.
Strategic Significance & Outlook
This hybrid quantum-MLIP approach provides a practical and scalable strategy for incorporating the unique advantages of quantum computing into mainstream materials R&D. It opens new avenues for the inverse design of novel materials, accelerated discovery of advanced catalysts, and precise engineering of functional surfaces. Industries such as energy (e.g., hydrogen storage, battery materials), pharmaceuticals (e.g., drug-surface interactions), and aerospace (e.g., high-temperature alloys) stand to benefit from the enhanced predictive capabilities. As quantum computing hardware continues to advance, this framework is poised to become an increasingly powerful tool, driving unprecedented innovation in material informatics and accelerating the transition from theoretical discovery to industrial application.
Source: https://papers.cool/arxiv/2609.21536
Get our weekly technology intelligence — free
Receive an infographic that lets you judge at a glance whether each field’s analysis report is worth reading.
Subscribe Free — Weekly Tech Intelligence
By subscribing, you’ll receive Troy-Technical’s weekly technology intelligence newsletter.
- Your email and selected fields are used only to deliver the newsletter.
- We never share your information with third parties.
- You can unsubscribe anytime via the link in each email.
See our Privacy Policy for details.
Takes about a minute · Unsubscribe anytime

Comments