TECHNOLOGY EXPLAINER
Fundamentals of Density Functional Theory
What a method that computes properties from the electron "density" can and cannot tell you
Papers and databases on materials calculations are full of numbers labelled "computed with DFT". DFT (density functional theory) is the most widely used method for computing energies, structures and band structures from the behaviour of a material's electrons. But computed values contain approximations and need not agree with measurements. This article uses figures to explain the idea of DFT, and sets out, from a materials engineer's viewpoint, what it can and cannot tell you, typical errors caused by the approximation (the functional) such as the underestimation of band gaps, how to read calculation settings, and the cost and reliability of the calculations. Electron correlation itself and the link to quantum computers are covered in our explainer on quantum chemistry and materials simulation, so here we concentrate on what is specific to DFT.
- The short version, what to know first, and where this fits in the series
- What DFT computes
- What it can and cannot tell you
- The ladder of approximations (functionals)
- Typical errors
- How to read the calculation settings
- Cost and scale
- Reproducibility and reliability
- A materials engineer's view
- Limits, what we could not confirm, and a summary
- How to read the yardsticks and standards
- Glossary
- References
- Claim-to-source audit
Sourced = stated in a published source (linked)
Our calculation = a value computed in this article from the stated assumptions
Not yet confirmed = plans, targets, drafts, work in progress, outlooks: not settled
Our own reading for materials engineers is marked as commentary. Inference is shown in bold. Computed values are stated together with the conditions of calculation (functional, settings), and the difference from measurement is kept separate.
1. The short version, what to know first, and where this fits in the series
- DFT is a calculation method based on the theorem that the ground-state energy is fixed by the "electron density" alone. Instead of following many electrons one by one, it repeatedly refines the electron density and outputs energies, forces and band structures Sourced[Refs. 1, 2]
- The theory is exact, but real calculations contain an approximation: the "functional". A typical error is the underestimation of band gaps. In a study comparing 472 materials with experiment, the mean error of the PBE gap was about −1.0 eV (mean absolute error 1.1 eV), against −0.1 eV (0.5 eV) for the hybrid functional HSE06 Sourced[Ref. 3]
- A computed value is a "value under conditions". A different functional, pseudopotential, cutoff, k-point set or DFT+U value changes the result. Even when the same structures were computed in three large databases, the median difference in band gap was about 9%, and the verdict "metal or not" disagreed for up to 7% of records Sourced[Ref. 4]
In the series' common analogy, DFT is "a physics simulation that computes properties from the behaviour of electrons" (it contains approximations and need not agree with measurement). In the division of labour in which calculation produces candidates cheaply and quickly, experiment confirms whether they are real, and AI narrows where to look, DFT is the "calculation" part. The "eV" (electronvolt) is a unit of energy, 1 eV ≈ 96.5 kJ/mol (the standard conversion). The "ground state" is the state of lowest energy.
A good order is our explainers on quantum chemistry and materials simulation (electron correlation and the limits of exact calculation) and on materials databases and the FAIR principles (where computed data are kept), then this article (how to read DFT itself). It leads on to our explainers on high-throughput computation (running calculations in large numbers), machine-learning interatomic potentials (learning a substitute for DFT) and graph neural networks (using DFT results as training data).
2. What DFT computes
The properties of matter are set by the behaviour of electrons. But with many electrons, following them one by one becomes very heavy (the reason is explained in our explainer on quantum chemistry and materials simulation). DFT changes the way of thinking.
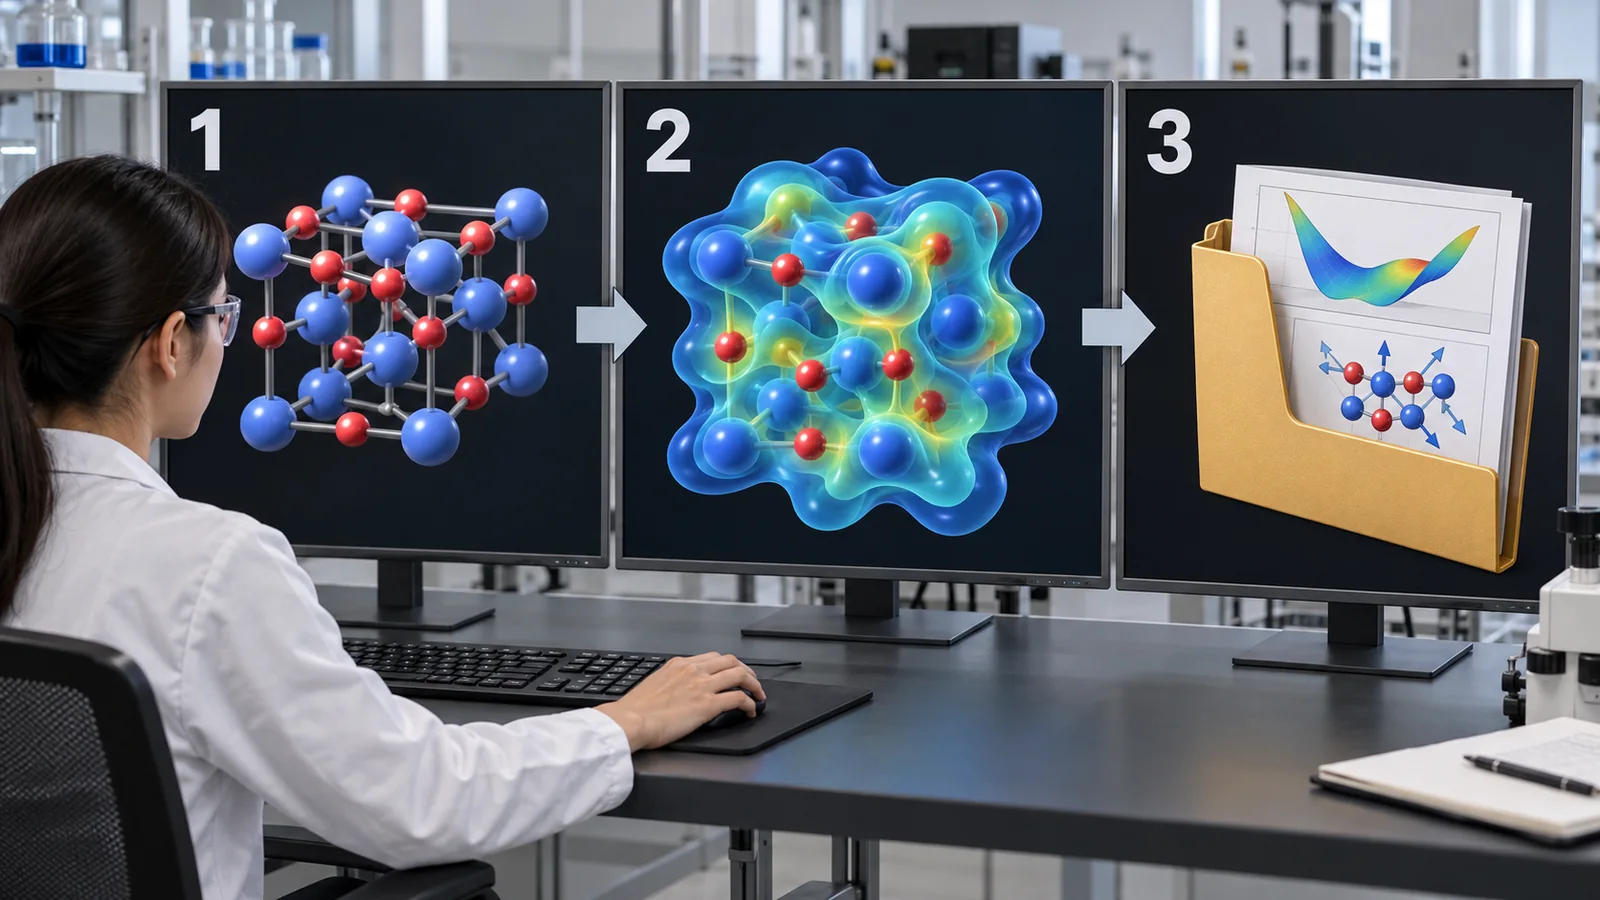
- 1964 (Hohenberg and Kohn): the ground-state energy is fully determined by the electron density (how many electrons there are at each point in space) Sourced[Ref. 1]. The theorem says that, without following electrons one by one, a "map of the density" is enough Sourced[Ref. 5].
- 1965 (Kohn and Sham): they showed how to replace the real problem of "interacting electrons" with one in which "non-interacting electrons move in an effective potential" Sourced[Ref. 6]. The potential is determined by the density.
View original SVG, values and sources
Original SVG reference: Concept diagram (vector drawing). The flow of a DFT calculation: guess a density, build the potential, solve the equations, and repeat until the new density equals the previous one.
In this scheme the potential is set by the density, and the density is set by the states of the electrons in the potential. So the calculation guesses a density, builds the potential, solves the equations and builds a new density, repeating until the density before and after is the same (self-consistent). This is the "self-consistent field (SCF) calculation". Kohn's Nobel lecture presents the framework in the order: the density as the basic variable, the Hohenberg-Kohn variational principle, and the self-consistent Kohn-Sham equations Sourced[Ref. 2].
The theorem guarantees that "a correct functional (a formula giving the energy from the density) exists". But its exact form is not known. In practice the formula is approximated (LDA, GGA and so on). The main cause of error is this approximation. This is the point of the "limit of the analogy" note at the bottom of the figure, and the next chapter, on the ladder of approximations, goes into it.
3. What it can and cannot tell you
To use DFT results in a materials decision, it matters to know first what it can and cannot tell you.
| What you want to know | From DFT | Explanation |
|---|---|---|
| Stable structure, bond strength | Yes | Structures can be optimised from the energy and forces of an atomic arrangement (the basic use of the Kohn-Sham framework) |
| Formation energy, stability | Mostly (mind the error) | In the OQMD check, the mean absolute error against 1,670 experimental formation energies was 0.096 eV/atom. Experiments also disagree among themselves by 0.082 eV/atom, and the authors suggest a share of the error comes from experimental uncertainty Sourced[Ref. 7] |
| Phonons (lattice vibrations) | Yes | Methods to compute phonons and related properties of crystals with density-functional perturbation theory are reviewed Sourced[Ref. 8] |
| Barriers for reactions and diffusion | Yes (movement of atoms) | The NEB method finds minimum-energy paths and saddle points; applications with plane-wave DFT are shown Sourced[Ref. 9] |
| Band gap | Needs care | Underestimated by many functionals (next chapter) |
| Excited states, optical absorption | Weak in the basic form | First-principles calculation of excited states needs more effort than ground-state DFT, and methods such as the GW approximation and the Bethe-Salpeter equation are used Sourced[Ref. 10] |
| Finite temperature, long times, microstructure, process | Not directly | DFT centres on ground-state calculations for hundreds to about a thousand atoms. The real performance of a material is often set by microstructure, impurities and process (commentary) |
The arrangement of the table is Our calculation (organisation). Checked on 23 October 2026.
"Stable" in DFT is a comparison at 0 K within that composition (the Materials Project glossary also explains that the convex hull has no temperature axis and is a 0 K prediction Sourced[Ref. 11]). Whether it can be synthesised also depends on kinetics (how fast reactions go), impurities and synthesis conditions. Calculation produces candidates; experiment confirms whether they can really be made.
4. The ladder of approximations (functionals)
What sets the accuracy of a real DFT calculation is the choice of the exchange-correlation functional (the approximation of the hard part of the energy as a function of the density). Perdew likened the approximations to "Jacob's ladder". Low rungs use only the density at each point, and higher rungs add more complex quantities built from the density (its gradient, the orbitals) and satisfy more of the exact conditions Sourced[Ref. 12].
- LDA (local density approximation): the simplest approximation, using only the density at each point (explained in the original Kohn-Sham paper and in Kohn's Nobel lecture) Sourced[Ref. 2].
- GGA (PBE and others): also uses the gradient of the density. PBE was derived as a simple GGA whose parameters are fundamental constants Sourced[Ref. 13]. It is one of the most widely used functionals in materials databases (the standard Materials Project calculation is PBE (GGA) Sourced[Ref. 14]).
- meta-GGA (SCAN, r²SCAN): also uses quantities built from the orbitals. SCAN is the first meta-GGA that obeys all 17 known exact constraints that a meta-GGA can, and is said to be remarkably accurate for lattice constants and weak interactions Sourced[Ref. 15]. r²SCAN restores constraint adherence to rSCAN, a version regularised for numerical performance Sourced[Ref. 16].
- Hybrids (HSE06, PBE0 and others): mix in part of the exact (Hartree-Fock) exchange. Because exact exchange is expensive in solids and metals, HSE uses a screened Coulomb potential to avoid this burden Sourced[Ref. 17].
There is no functional that is "always best". Haas et al. found that for lattice constants of solids, newer GGA-type functionals often improve on LDA and PBE, but "none of them can be considered as good for all" the solids they investigated Sourced[Ref. 18]. Mardirossian and Head-Gordon assessed 200 functionals on a molecular database (nearly 5,000 data points) and gave recommendations, but those are results for molecules Sourced[Ref. 19]. Medvedev et al. point out that some recent functionals, while better in energies, have gone backwards in the correctness of the electron density since the early 2000s Sourced[Ref. 1]. Choose a functional that has been tested for the property and the type of material you need (commentary).
A different functional gives different values even for the same material. So values should be compared between the same functional and settings. For mixing PBE (GGA), DFT+U and r²SCAN calculations, the Materials Project provides a way to correct the energies and place them on a single phase diagram Sourced[Ref. 20]. This is the flip side of the fact that values from different functionals cannot be compared as they are.
5. Typical errors
(1) Underestimated band gaps
The best-known error of DFT is the underestimation of band gaps. Perdew and Levy (1983) argued that the local-density approximation underestimates the fundamental gaps of semiconductors and insulators by about 40%, and that even the exact Kohn-Sham theory is expected to underestimate the gap, because of the derivative discontinuity of the exchange-correlation energy Sourced[Ref. 21]. Sham and Schlüter showed that the true gap includes a correction from this discontinuity Sourced[Ref. 22].
The Materials Project documentation explains that the correspondence between Kohn-Sham energies and real electron energies is theoretically valid only for the highest occupied state. It says LDA and GGA gaps are about 50% too small in the literature, and in its own internal test (237 compounds) about a factor of 1.6 too small on average, with a mean absolute error of 0.6 eV even after a constant shift Sourced[Ref. 23].
- Borlido et al. compared 12 functionals on 472 nonmagnetic materials with known experimental structure and gap. Some results (rounded): for PBE the mean error of the gap is about −1.0 eV and the mean absolute error 1.1 eV; for HSE06 −0.1 eV and 0.5 eV; for mBJ −0.2 eV and 0.5 eV Sourced[Ref. 3].
- PBE wrongly computed 31 of the 472 materials (6.6%) as metals (gap below 0.01 eV). For HSE06 the number was 10 Sourced[Ref. 3].
- mBJ is a potential designed for band gaps, and its purpose differs from that of the other functionals. The study concluded that mBJ is at present the best, closely followed by HSE06 Sourced[Ref. 3].
- Perdew et al. explain why, in generalized Kohn-Sham theory, the band gaps of meta-GGAs and hybrids can be more realistic than those of GGAs or of the exact Kohn-Sham potential Sourced[Ref. 24].
The gap from ordinary DFT (PBE and the like) cannot stand in for the measured gap. It is systematically small, and semiconductors and insulators sometimes come out as "metals". The shape of the band structure (band dispersion) is said to agree with experiment within about 0.1 to 0.4 eV in one small test, but the gap is a different matter. If you use the gap, state which of the following you did: (1) use a higher approximation such as a hybrid functional; (2) fix a constant shift (a scissor correction) by comparison with experiment; (3) use only relative trends (the order among materials) Sourced[Ref. 23].
(2) Transition-metal oxides: self-interaction error and DFT+U
For d electrons and for oxides, GGA has another error. Perdew and Zunger (1981) pointed out that approximate functionals carry a "self-interaction" in which an electron repels itself, which the exact functional does not, and proposed a correction Sourced[Ref. 25]. Wang et al. identified two substantial errors in the oxidation energies of transition-metal oxides. One is the overbinding of the O₂ molecule in GGA (a constant error that occurs only when the oxidant is O₂); the other is the correlation error in 3d orbitals (self-interaction penalises a reduced state). The first can be corrected by fitting the formation enthalpies of simple oxides, and the second can be handled by GGA+U; they report that oxidation energies then agree well with experiment for reasonable and consistent values of U Sourced[Ref. 26].
DFT+U adds a Hubbard U (a parameter for the repulsion between two electrons in the same orbital) to d electrons and the like; Anisimov et al. used it for Mott insulators and Dudarev et al. for the electron energy-loss spectra and structural stability of NiO Sourced[Refs. 27, 28]. In the Materials Project, oxides and fluorides of Co, Cr, Fe, Mn, Mo, Ni, V and W use values of U fitted to experimental formation enthalpies (U for V is 3.25 eV) Sourced[Ref. 29]. Jain et al. report that a method that mixes GGA and GGA+U total energies reduced the mean absolute relative error of formation enthalpies of 49 ternary oxides from about 7.7-21% in GGA+U to under 2% Sourced[Ref. 30].
The U of the Materials Project is a value decided by fitting to experimental data, calibrated for each system (oxides and fluorides) Sourced[Ref. 29]. A calculation using U is therefore "a calculation tuned to agree for that kind of compound with that value of U". It does not necessarily carry over to other properties (gaps, magnetism) or other kinds of compound (such as sulfides) (commentary).
(3) Other typical cases
- van der Waals forces: standard Kohn-Sham DFT does not include them. The DFT-D method of Grimme et al. adds a dispersion correction, usable for 94 elements, as an extra term Sourced[Ref. 31]. In layered materials and molecular crystals, whether the correction is used can decide the result.
- Lattice constants: PBE is said to give excellent results for the structure of solids in many cases. Among other GGA-type functionals, none is good for all solids Sourced[Ref. 18].
- Differences between databases: covered in the chapter on reproducibility and reliability below.
6. How to read the calculation settings
When you read computed values in a paper or a database, always check the "calculation conditions". The main items and notes on reading them are in the table below.
| Setting | Meaning | Reading note |
|---|---|---|
| Functional | The approximation of the exchange-correlation energy (LDA, PBE, SCAN, HSE06 and so on) | Do not compare values from different functionals as they are; their error tendencies differ |
| DFT+U and the value of U | An adjustment that compensates for the self-interaction error of d and f electrons | Which elements, and which U? What was it tuned for (oxides and so on)? |
| Pseudopotential, PAW | An approximation that replaces core electrons with something easier to compute (PAW is Blöchl's method) Sourced[Ref. 32] | Several versions exist for each element. In some cases the choice changes the thermodynamic result |
| Cutoff energy | The size of the plane-wave basis | Too low means unconverged. Raise it until the result is within the tolerance for the property you need |
| k-points | The fineness of the sampling of the Brillouin zone (the Monkhorst-Pack method and others) Sourced[Ref. 33] | Finer for metals. A larger cell needs fewer |
| Convergence criteria | The conditions for ending the SCF and structure optimisation | Tighter for calculations that need a precise structure, such as elastic constants and phonons |
| Spin and magnetic arrangement | Whether there is magnetism and the direction of the moments | The choice of magnetic order changes the energy. Databases can disagree on whether a material is magnetic |
| Cell size | Defects, interfaces and surfaces are represented in large periodic cells | If the cell is small, defects interact with their own images |
| Software and version | VASP, Quantum ESPRESSO and so on | Even with the same settings, differences in the implementation show up (next chapter) |
The arrangement of the table is Our calculation (organisation). Commentary based on primary sources such as the Materials Project documentation. Checked on 23 October 2026.
Example: the Materials Project settings
- Standard calculations (PBE/PBE+U): the PAW method; a plane-wave cutoff of 520 eV (1.3 times the highest cutoff recommended among the pseudopotentials used); a baseline k-point mesh of 1000 divided by the number of atoms in the cell; the Monkhorst-Pack method (Γ-centred for hexagonal cells); the tetrahedron method for integration; and an ionic convergence energy difference of 0.0005 × (number of atoms) per cell Sourced[Ref. 14].
- Checks of convergence: with an older setting (500 k-points per reciprocal atom), most compounds tested converged in total energy to within 5 meV/atom and 96% to within 15 meV/atom, and oxides were generally below 1 meV/atom Sourced[Ref. 14].
- r²SCAN calculations: a cutoff of 680 eV and k-point grids from KSPACING values of 0.22 to 0.44 Å⁻¹ (decided from the GGA-estimated gap), chosen so that formation energies converge to about 1 meV/atom on a benchmark set of 21 materials Sourced[Ref. 34].
- Current state (date checked): the latest Materials Project database version is v2026.09.28 (live on 19 October 2026), and recomputation of all materials with r²SCAN is under way (about 26% of non-GNoME materials in the April 2026 entry; this recomputed data was not yet integrated into the core materials data) Sourced[Ref. 35].
It means that even if you raise the settings (cutoff, k-points), the property you need does not change by more than you can tolerate. Energy differences (formation energies, barriers) often converge before the total energy itself, so check with the differences. If the target is the gap or phonons, check with that property (commentary).
7. Cost and scale
The cost of DFT rises steeply with the number of atoms. The review by Bowler and Miyazaki explains that the cost of standard methods grows as the cube of the number of atoms (linear-scaling methods grow in proportion to it) and becomes steep at around a thousand atoms. It also notes that orthogonalisation becomes the most expensive step at a few hundred atoms Sourced[Ref. 36].
- Under the cube-law assumption, a tenfold increase in atoms (10 to 100) means 1,000 times the cost, and a hundredfold increase (10 to 1000) a million times Our calculation.
- Making a 2×2×2 supercell of a crystal multiplies the number of atoms by 8, but the k-points drop from 8×8×8 (512) to 4×4×4 (64). Assuming cost ∝ (number of k-points) × (number of atoms)³, the ratio is (64 / 512) × (32 / 4)³ = 64 times Our calculation. This is why calculations for defects, interfaces and surfaces (which need large cells) are heavy.
- This count rests on the formal scaling of the standard method. Actual run times vary with the code, the parallelisation and the material (metal or insulator).
8. Reproducibility and reliability
"If someone else computes the same material with different software, do they get the same value?" is an important question for a materials engineer.
- Agreement between codes (verification): a community study by Lejaeghere et al. compared PBE equations of state for 71 elemental crystals across 15 solid-state codes and 40 potentials or basis types, and showed that recent codes and pseudopotentials agree very well, with differences comparable to those between high-precision experiments, while older methods agree less well Sourced[Ref. 37]. The Expert Recommendation by Bosoni et al. proposes extending this verification to 960 equations of state covering hydrogen to curium (Z = 1 to 96) Sourced[Ref. 38]. Prandini et al. verified pseudopotential libraries by comparison with all-electron calculations and other tests, and gave sets for high-throughput work (efficiency) and for high precision (SSSP) Sourced[Ref. 39].
- Agreement between databases: Hegde et al. compared calculations in AFLOW, the Materials Project and OQMD starting from the same crystal structures, and showed that formation energies and volumes agree fairly well, while band gaps and magnetic moments agree less well. The verdict on metal versus insulator disagreed for up to 7% of records and that on magnetism for up to 15%, and the size of the differences was comparable to the difference between DFT and experiment. They trace causes to the choice of pseudopotentials, DFT+U and the elemental reference states Sourced[Ref. 4].
- Difference from experiment: in the OQMD check, the difference between DFT and experiment for formation energy (0.096 eV/atom) is close to that between experiments themselves (0.082 eV/atom), and the authors suggest part of the error comes from experimental uncertainty Sourced[Ref. 7].
(1) Look at differences under the same conditions rather than absolute values (material A against material B, or structure A against structure B, with the same functional and settings). (2) Compare the size of the error with the width of the decision (for example, if the formation-energy error is about 0.1 eV/atom, do not call something stable or unstable on the strength of a smaller difference). (3) Read gaps and magnetism qualitatively (they are the quantities that even databases disagree on). Do not write a DFT value as if it were a measured one; write it as "computed (PBE, software X)" to keep them apart (commentary).
9. A materials engineer's view
The questions DFT is good at are energy comparisons at the atomic scale: "which of structure A and structure B is more stable?", "how does the bonding change if we add this element?", "how readily does this defect form?", "how easily do ions move along this path?". It is weak at questions in which microstructure, process and time matter, such as "how many years will this material last at this temperature?". Framing a question for calculation as (1) recast it as an atomic-scale comparison and (2) link it to a quantity you can measure (phase identification, capacity, diffusion coefficient) makes DFT easier to put to use (commentary).
For a candidate that calculation calls "stable" or "with a suitable gap", check in turn: (1) was the target phase obtained (telling it from known compounds, for example by powder X-ray diffraction); (2) does it reproduce (another lot); (3) the effect of impurities and disorder; (4) was it compared with a control (a known material) by the same measurement. Recording the calculation conditions (the functional and so on) and the difference from the measurement helps in judging how far to trust the next calculation (commentary).
10. Limits, what we could not confirm, and a summary
(1) What we could not confirm
- We limited ourselves to a summary of the main functionals (LDA, PBE, SCAN, mBJ, HSE06, PBE0) and did not give the figures for all 12 functionals of Borlido et al., nor the errors per material.
- For Haas et al. we stay within the abstract and did not confirm the specific numerical errors in lattice constants.
- For the Materials Project we stay with the entry closest to the check date (April 2026) and did not confirm the latest progress of the recomputation.
- The "cube" of the cost is a formal estimate; we did not confirm measured run times for each code.
- We could not confirm an international standard (ISO/IEC or similar) that defines evaluation or reproducibility of DFT results.
- DFT rests on the theorem that the electron density fixes the ground-state energy, and computes by repeatedly refining the density. The theory is exact, but the approximation of the functional is the main cause of error Sourced
- What it tells you: structures, bonding, formation energies, phonons, barriers. What needs care: band gaps, excitations, strong correlation, van der Waals forces. What it does not answer directly: microstructure, process, time. Sourced
- Band gaps are systematically small with PBE: a mean error of about −1.0 eV over 472 materials, with 31 false metals. Hybrids and mBJ tend to have smaller errors Sourced
- A computed value is a value under conditions. Check the functional, DFT+U, pseudopotential, cutoff and k-points, and compare only like with like. Even databases disagree on gaps and magnetism Sourced
- Cost grows roughly as the cube of the number of atoms. Defects and interfaces need large cells and are heavy Our calculation
11. How to read the yardsticks and standards
The first yardstick of DFT is the difference from experiment (formation energy, lattice constant, band gap). Next come agreement between codes and databases (verification) and convergence (does the result stay put when the settings are raised). Read each as a yardstick for something different: the difference from experiment is the "error of the method", and the difference between codes is the "precision of the implementation". Checked on 23 October 2026.
| Item | Representative benchmarks and methods (name, source) | Standards and guidance (ISO/IEC, NIST, etc.) | Reading notes |
|---|---|---|---|
| Agreement between codes (verification) | Comparison of equations of state of 71 elemental crystals with the Δ gauge and others (Lejaeghere et al. 2016) [Ref. 37]Sourced Verification protocol with 960 equations of state (Bosoni et al. 2024) [Ref. 38]Sourced Verification of pseudopotentials (SSSP, Prandini et al. 2018) [Ref. 39]Sourced | Not found (we did not find an international standard for verifying DFT implementations) | Verification on elemental crystals (PBE). It does not necessarily carry over to compounds or other functionals |
| Accuracy of functionals | Band gaps: 472 materials (Borlido et al. 2019) [Ref. 3]. Lattice constants of solids (Haas et al. 2009) [Ref. 18]. Molecules: nearly 5,000 data points and 200 functionals (Mardirossian and Head-Gordon 2017) [Ref. 19]Sourced | Not found | Results limited to the kind of material in the benchmark (molecules, solids, nonmagnetic and so on) |
| Formation energy (against experiment) | OQMD: comparison with 1,670 experimental formation energies (Kirklin et al. 2015) [Ref. 7]Sourced | Not found | Experiments themselves scatter (0.082 eV/atom) |
| Agreement between databases | Comparison of AFLOW, the Materials Project and OQMD (Hegde et al. 2023) [Ref. 4]Sourced | Not found | Gaps and magnetism tend to disagree. The authors say standardising the calculation settings would help |
| Overall comparison of methods | JARVIS-Leaderboard: a framework that compares electronic-structure methods, software and pseudopotentials with experiment (NIST; Choudhary et al. 2024) [Ref. 40]Sourced | Not found | A public venue for comparison. Check whether it covers your own material system |
The arrangement of the table is Our calculation (organisation). Checked in the papers and official pages (23 October 2026). We did not find an international standard specific to DFT, so the standards column says "Not found".
Reading notes (for materials engineers)
- A benchmark ranking is not the same as strength on your own materials: the materials in Borlido et al. are nonmagnetic materials with known experimental structures. Magnetic materials, defects and interfaces need separate checks [Ref. 3]
- Verification (agreement between codes) and validity (agreement with experiment) are different: what Lejaeghere et al. showed is agreement between codes, not that PBE agrees with experiment [Ref. 37]
- Distinguish plans from achievements: the r²SCAN recomputation in the Materials Project is under way and integration of the data is partial (Not yet confirmed) [Ref. 35]
12. Glossary
- DFT (density functional theory)
- A theory in which the electron density fixes the ground-state energy, and the calculation methods based on it.
- Electron density n(r)
- The distribution of how many electrons there are at each point in space.
- Ground state
- The state of lowest energy.
- Kohn-Sham equations
- Equations in which non-interacting electrons move in an effective potential, replacing the real problem.
- SCF (self-consistent field)
- A calculation that refines the density and the potential repeatedly until they no longer disagree.
- Exchange-correlation functional
- An approximate formula that evaluates the complicated effects between electrons from the density. The main source of error.
- LDA / GGA / meta-GGA / hybrid
- Rungs of functional approximation; in order, more ingredients are used.
- PBE
- A widely used GGA functional; the standard in materials databases.
- SCAN / r²SCAN
- Meta-GGA functionals built to satisfy exact constraints.
- HSE06
- A hybrid functional that uses a screened Coulomb interaction; suited to solids.
- Band gap
- The energy gap between filled and empty electron states.
- False metal
- The error of computing a gap of zero for what is really a semiconductor or insulator.
- Self-interaction error
- An approximation error in which an electron repels itself; conspicuous for d electrons.
- DFT+U
- A method that adds a Hubbard U to d and f electrons to compensate for the self-interaction error.
- Pseudopotential / PAW
- Approximations that replace core electrons with something easier to compute.
- Cutoff energy
- The value that sets the size of the plane-wave basis.
- k-points
- Points in the Brillouin zone used for integration. More points means finer sampling.
- Convergence
- The result no longer changes beyond tolerance when the settings are raised.
- Supercell
- A large cell made by repeating the crystal's cell several times; used for defects.
- Phonon
- The quantum of lattice vibration in a crystal.
- NEB method
- A method for finding the minimum-energy path of a reaction or diffusion.
- GW approximation
- A method that gives excited states (such as band gaps) more accurately than DFT.
- Verification
- Checking whether different codes give the same answer to the same problem.
13. References
- Medvedev M. G. et al. "Density functional theory is straying from the path toward the exact functional", Science 355, 49–52 (2017), DOI 10.1126/science.aah5975 — doi.org/10.1126/science.aah5975
- Kohn W. "Nobel lecture: electronic structure of matter — wave functions and density functionals", Rev. Mod. Phys. 71, 1253–1266 (1999), DOI 10.1103/RevModPhys.71.1253 — doi.org/10.1103/RevModPhys.71.1253
- Borlido P. et al. "Large-scale benchmark of exchange-correlation functionals for the determination of electronic band gaps of solids", J. Chem. Theory Comput. 15, 5069–5079 (2019), DOI 10.1021/acs.jctc.9b00322 — doi.org/10.1021/acs.jctc.9b00322
- Hegde V. I. et al. "Quantifying uncertainty in high-throughput density functional theory: a comparison of AFLOW, Materials Project, and OQMD", Phys. Rev. Mater. 7, 053805 (2023), DOI 10.1103/PhysRevMaterials.7.053805 — doi.org/10.1103/PhysRevMaterials.7.053805
- Hohenberg P., Kohn W. "Inhomogeneous electron gas", Phys. Rev. 136, B864–B871 (1964), DOI 10.1103/PhysRev.136.B864 — doi.org/10.1103/PhysRev.136.B864
- Kohn W., Sham L. J. "Self-consistent equations including exchange and correlation effects", Phys. Rev. 140, A1133–A1138 (1965), DOI 10.1103/PhysRev.140.A1133 — doi.org/10.1103/PhysRev.140.A1133
- Kirklin S. et al. "The Open Quantum Materials Database (OQMD): assessing the accuracy of DFT formation energies", npj Comput. Mater. 1, 15010 (2015), DOI 10.1038/npjcompumats.2015.10 — doi.org/10.1038/npjcompumats.2015.10
- Baroni S. et al. "Phonons and related crystal properties from density-functional perturbation theory", Rev. Mod. Phys. 73, 515–562 (2001), DOI 10.1103/RevModPhys.73.515 — doi.org/10.1103/RevModPhys.73.515
- Henkelman G., Uberuaga B. P., Jónsson H. "A climbing image nudged elastic band method for finding saddle points and minimum energy paths", J. Chem. Phys. 113, 9901–9904 (2000), DOI 10.1063/1.1329672 — doi.org/10.1063/1.1329672
- Onida G., Reining L., Rubio A. "Electronic excitations: density-functional versus many-body Green's-function approaches", Rev. Mod. Phys. 74, 601–659 (2002), DOI 10.1103/RevModPhys.74.601 — doi.org/10.1103/RevModPhys.74.601
- Materials Project "Documentation: Glossary of terms (energy above hull)", (Materials Project documentation) — docs.materialsproject.org/frequently-asked-questions/glossary-of-terms
- Perdew J. P., Schmidt K. "Jacob's ladder of density functional approximations for the exchange-correlation energy", AIP Conf. Proc. 577, 1–20 (2001), DOI 10.1063/1.1390175 — doi.org/10.1063/1.1390175
- Perdew J. P., Burke K., Ernzerhof M. "Generalized gradient approximation made simple", Phys. Rev. Lett. 77, 3865–3868 (1996), DOI 10.1103/PhysRevLett.77.3865 — doi.org/10.1103/PhysRevLett.77.3865
- Materials Project "Documentation: GGA/GGA+U calculations — Parameters and convergence", (Materials Project documentation) — docs.materialsproject.org/methodology/materials-methodology/calculation-details/gga+u-calculations/parameters-and-convergence
- Sun J., Ruzsinszky A., Perdew J. P. "Strongly constrained and appropriately normed semilocal density functional", Phys. Rev. Lett. 115, 036402 (2015), DOI 10.1103/PhysRevLett.115.036402 — doi.org/10.1103/PhysRevLett.115.036402
- Furness J. W. et al. "Accurate and numerically efficient r²SCAN meta-generalized gradient approximation", J. Phys. Chem. Lett. 11, 8208–8215 (2020), DOI 10.1021/acs.jpclett.0c02405 — doi.org/10.1021/acs.jpclett.0c02405
- Heyd J., Scuseria G. E., Ernzerhof M. "Hybrid functionals based on a screened Coulomb potential", J. Chem. Phys. 118, 8207–8215 (2003), DOI 10.1063/1.1564060 — doi.org/10.1063/1.1564060
- Haas P., Tran F., Blaha P. "Calculation of the lattice constant of solids with semilocal functionals", Phys. Rev. B 79, 085104 (2009), DOI 10.1103/PhysRevB.79.085104 — doi.org/10.1103/PhysRevB.79.085104
- Mardirossian N., Head-Gordon M. "Thirty years of density functional theory in computational chemistry: an overview and extensive assessment of 200 density functionals", Mol. Phys. 115, 2315–2372 (2017), DOI 10.1080/00268976.2017.1333644 — doi.org/10.1080/00268976.2017.1333644
- Materials Project "Documentation: GGA/GGA+U/r2SCAN mixing (energy corrections)", (Materials Project documentation) — docs.materialsproject.org/methodology/materials-methodology/thermodynamic-stability/thermodynamic-stability/gga-gga+u-r2scan-mixing
- Perdew J. P., Levy M. "Physical content of the exact Kohn-Sham orbital energies: band gaps and derivative discontinuities", Phys. Rev. Lett. 51, 1884–1887 (1983), DOI 10.1103/PhysRevLett.51.1884 — doi.org/10.1103/PhysRevLett.51.1884
- Sham L. J., Schlüter M. "Density-functional theory of the energy gap", Phys. Rev. Lett. 51, 1888–1891 (1983), DOI 10.1103/PhysRevLett.51.1888 — doi.org/10.1103/PhysRevLett.51.1888
- Materials Project "Documentation: Electronic Structure (accuracy of band structures and band gaps)", (Materials Project documentation) — docs.materialsproject.org/methodology/materials-methodology/electronic-structure
- Perdew J. P. et al. "Understanding band gaps of solids in generalized Kohn-Sham theory", Proc. Natl. Acad. Sci. USA 114, 2801–2806 (2017), DOI 10.1073/pnas.1621352114 — doi.org/10.1073/pnas.1621352114
- Perdew J. P., Zunger A. "Self-interaction correction to density-functional approximations for many-electron systems", Phys. Rev. B 23, 5048–5079 (1981), DOI 10.1103/PhysRevB.23.5048 — doi.org/10.1103/PhysRevB.23.5048
- Wang L., Maxisch T., Ceder G. "Oxidation energies of transition metal oxides within the GGA+U framework", Phys. Rev. B 73, 195107 (2006), DOI 10.1103/PhysRevB.73.195107 — doi.org/10.1103/PhysRevB.73.195107
- Anisimov V. I., Zaanen J., Andersen O. K. "Band theory and Mott insulators: Hubbard U instead of Stoner I", Phys. Rev. B 44, 943–954 (1991), DOI 10.1103/PhysRevB.44.943 — doi.org/10.1103/PhysRevB.44.943
- Dudarev S. L. et al. "Electron-energy-loss spectra and the structural stability of nickel oxide: an LSDA+U study", Phys. Rev. B 57, 1505–1509 (1998), DOI 10.1103/PhysRevB.57.1505 — doi.org/10.1103/PhysRevB.57.1505
- Materials Project "Documentation: Hubbard U values", (Materials Project documentation) — docs.materialsproject.org/methodology/materials-methodology/calculation-details/gga+u-calculations/hubbard-u-values
- Jain A. et al. "Formation enthalpies by mixing GGA and GGA+U calculations", Phys. Rev. B 84, 045115 (2011), DOI 10.1103/PhysRevB.84.045115 — doi.org/10.1103/PhysRevB.84.045115
- Grimme S. et al. "A consistent and accurate ab initio parametrization of density functional dispersion correction (DFT-D) for the 94 elements H-Pu", J. Chem. Phys. 132, 154104 (2010), DOI 10.1063/1.3382344 — doi.org/10.1063/1.3382344
- Blöchl P. E. "Projector augmented-wave method", Phys. Rev. B 50, 17953–17979 (1994), DOI 10.1103/PhysRevB.50.17953 — doi.org/10.1103/PhysRevB.50.17953
- Monkhorst H. J., Pack J. D. "Special points for Brillouin-zone integrations", Phys. Rev. B 13, 5188–5192 (1976), DOI 10.1103/PhysRevB.13.5188 — doi.org/10.1103/PhysRevB.13.5188
- Materials Project "Documentation: r2SCAN calculations — Parameters and convergence", (Materials Project documentation) — docs.materialsproject.org/methodology/materials-methodology/calculation-details/r2scan-calculations/parameters-and-convergence
- Materials Project "Documentation: Database versions (v2026.09.28 and v2026.04.13 entries; page read on 23 October 2026)", (Materials Project documentation) — docs.materialsproject.org/changes/database-versions
- Bowler D. R., Miyazaki T. "O(N) methods in electronic structure calculations", Rep. Prog. Phys. 75, 036503 (2012), DOI 10.1088/0034-4885/75/3/036503 — doi.org/10.1088/0034-4885/75/3/036503
- Lejaeghere K. et al. "Reproducibility in density functional theory calculations of solids", Science 351, aad3000 (2016), DOI 10.1126/science.aad3000 — doi.org/10.1126/science.aad3000
- Bosoni E. et al. "How to verify the precision of density-functional-theory implementations via reproducible and universal workflows", Nat. Rev. Phys. 6, 45–58 (2024), DOI 10.1038/s42254-023-00655-3 — doi.org/10.1038/s42254-023-00655-3
- Prandini G. et al. "Precision and efficiency in solid-state pseudopotential calculations", npj Comput. Mater. 4, 72 (2018), DOI 10.1038/s41524-018-0127-2 — doi.org/10.1038/s41524-018-0127-2
- Choudhary K. et al. "JARVIS-Leaderboard: a large scale benchmark of materials design methods", npj Comput. Mater. 10, 93 (2024), DOI 10.1038/s41524-024-01259-w — doi.org/10.1038/s41524-024-01259-w
14. Claim-to-source audit
| Claim in the text | Basis | Category |
|---|---|---|
| The ground-state energy being fully determined by the electron density (Hohenberg-Kohn); replacing the interacting problem with independent electrons in an effective potential (Kohn-Sham); the structure of Kohn's Nobel lecture (density as the basic variable, variational principle, self-consistent Kohn-Sham equations, LDA) | Reference 5 https://doi.org/10.1103/PhysRev.136.B864 Reference 6 https://doi.org/10.1103/PhysRev.140.A1133 Reference 2 https://doi.org/10.1103/RevModPhys.71.1253 Reference 1 https://doi.org/10.1126/science.aah5975 | Sourced |
| Jacob's ladder (higher rungs add more complex ingredients and can satisfy more exact conditions); PBE (a GGA whose parameters are fundamental constants); SCAN (the first meta-GGA to satisfy all 17 known exact constraints, with remarkable accuracy for lattice constants and weak interactions); r²SCAN (restores constraint adherence to rSCAN); HSE (a screened Coulomb potential avoids the cost of exact exchange); the point by Medvedev et al. that some functionals lost ground in electron-density correctness | Reference 12 https://doi.org/10.1063/1.1390175 Reference 13 https://doi.org/10.1103/PhysRevLett.77.3865 Reference 15 https://doi.org/10.1103/PhysRevLett.115.036402 Reference 16 https://doi.org/10.1021/acs.jpclett.0c02405 Reference 17 https://doi.org/10.1063/1.1564060 Reference 1 https://doi.org/10.1126/science.aah5975 | Sourced |
| Haas et al.: for lattice constants of solids newer GGAs often improve on LDA and PBE, but none is good for all solids; Mardirossian and Head-Gordon: 200 functionals assessed on a molecular database of nearly 5,000 data points | Reference 18 https://doi.org/10.1103/PhysRevB.79.085104 Reference 19 https://doi.org/10.1080/00268976.2017.1333644 | Sourced |
| Perdew and Levy: LDA underestimates gaps by about 40% and even exact Kohn-Sham is expected to underestimate them because of derivative discontinuities; Sham and Schlüter: the correction from the discontinuity; Perdew et al.: why meta-GGA and hybrid gaps can be more realistic in generalized Kohn-Sham theory | Reference 21 https://doi.org/10.1103/PhysRevLett.51.1884 Reference 22 https://doi.org/10.1103/PhysRevLett.51.1888 Reference 24 https://doi.org/10.1073/pnas.1621352114 | Sourced |
| Borlido et al.: 472 nonmagnetic materials and 12 functionals; mean errors and mean absolute errors for LDA, PBE, SCAN, mBJ, HSE06 and PBE0 (rounded: LDA -1.2 and 1.2, PBE -1.0 and 1.1, SCAN -0.7 and 0.8, mBJ -0.2 and 0.5, HSE06 -0.1 and 0.5, PBE0 +0.5 and 0.8 eV), numbers of false metals (35, 31, 19, 11, 10, 5), and the conclusion that mBJ is best with HSE06 close behind | Reference 3 https://doi.org/10.1021/acs.jctc.9b00322 | Sourced |
| Materials Project: the correspondence between Kohn-Sham eigenvalues and real energies is rigorous only for the highest occupied state; LDA/GGA gaps are about 50% too small in the literature and on average a factor of 1.6 too small in the internal test of 237 compounds, with MAE 0.6 eV even after a shift; band dispersion errors of 0.1-0.4 eV (a small test); the scissor operation | Reference 23 https://docs.materialsproject.org/methodology/materials-methodology/electronic-structure | Sourced |
| Perdew and Zunger: self-interaction; Wang et al.: O₂ overbinding and 3d correlation error, GGA+U; Anisimov et al. and Dudarev et al.: DFT+U; Materials Project: U values fitted to experimental formation enthalpies for oxides and fluorides of Co, Cr, Fe, Mn, Mo, Ni, V and W (3.25 eV for V); Jain et al.: error reduced from about 7.7-21% to under 2% for 49 ternary oxides | Reference 25 https://doi.org/10.1103/PhysRevB.23.5048 Reference 26 https://doi.org/10.1103/PhysRevB.73.195107 Reference 27 https://doi.org/10.1103/PhysRevB.44.943 Reference 28 https://doi.org/10.1103/PhysRevB.57.1505 Reference 29 https://docs.materialsproject.org/methodology/materials-methodology/calculation-details/gga+u-calculations/hubbard-u-values Reference 30 https://doi.org/10.1103/PhysRevB.84.045115 | Sourced |
| Grimme et al.'s DFT-D (94 elements); PAW (Blöchl); Monkhorst-Pack k-points; the Materials Project settings (PBE with PAW, 520 eV = 1.3 times the largest recommended value, k-points 1000 per atoms in the cell, tetrahedron method, convergence statements: most compounds within 5 meV/atom and 96% within 15 meV/atom for the older setting), r²SCAN (680 eV, KSPACING 0.22-0.44 Å⁻¹, about 1 meV/atom on 21 materials), and the mixing corrections between functionals | Reference 31 https://doi.org/10.1063/1.3382344 Reference 32 https://doi.org/10.1103/PhysRevB.50.17953 Reference 33 https://doi.org/10.1103/PhysRevB.13.5188 Reference 14 https://docs.materialsproject.org/methodology/materials-methodology/calculation-details/gga+u-calculations/parameters-and-convergence Reference 34 https://docs.materialsproject.org/methodology/materials-methodology/calculation-details/r2scan-calculations/parameters-and-convergence Reference 20 https://docs.materialsproject.org/methodology/materials-methodology/thermodynamic-stability/thermodynamic-stability/gga-gga+u-r2scan-mixing | Sourced |
| Materials Project database version v2026.09.28 (released 19 October 2026); the April 2026 entry (r²SCAN recomputation at about 26% of non-GNoME materials, not yet integrated into the core data); the glossary description of the convex hull (no temperature axis, a 0 K prediction) | Reference 35 https://docs.materialsproject.org/changes/database-versions Reference 11 https://docs.materialsproject.org/frequently-asked-questions/glossary-of-terms | Sourced |
| OQMD: comparison with 1,670 experimental formation energies (MAE 0.096 eV/atom) and the disagreement among experiments (0.082 eV/atom); phonons (density-functional perturbation theory), the NEB method, excited-state calculations (GW and Bethe-Salpeter); cost of standard methods growing as the cube of the number of atoms, steep at about a thousand atoms, with orthogonalisation the most expensive step at a few hundred | Reference 7 https://doi.org/10.1038/npjcompumats.2015.10 Reference 8 https://doi.org/10.1103/RevModPhys.73.515 Reference 9 https://doi.org/10.1063/1.1329672 Reference 10 https://doi.org/10.1103/RevModPhys.74.601 Reference 36 https://doi.org/10.1088/0034-4885/75/3/036503 | Sourced |
| Lejaeghere et al.: PBE equations of state of 71 elemental crystals across 15 codes and 40 potentials or basis types, recent codes agreeing to within differences comparable to those between high-precision experiments; Bosoni et al.: Z = 1 to 96, 960 equations of state; Prandini et al.: SSSP; Hegde et al.: comparison of AFLOW, MP and OQMD (formation energy up to 0.105 eV/atom and median 6%, volume 0.65 ų/atom and 4%, band gap 0.21 eV and 9%, magnetic moment 0.15 µB/f.u. and 8%, metal-versus-insulator disagreement up to 7%, magnetism disagreement up to 15%); JARVIS-Leaderboard | Reference 37 https://doi.org/10.1126/science.aad3000 Reference 38 https://doi.org/10.1038/s42254-023-00655-3 Reference 39 https://doi.org/10.1038/s41524-018-0127-2 Reference 4 https://doi.org/10.1103/PhysRevMaterials.7.053805 Reference 40 https://doi.org/10.1038/s41524-024-01259-w | Sourced |
| Atom counts and cost (assuming N³: 1,000 times from 10 to 100 atoms, a million times from 10 to 1000), and the supercell example (4 atoms with 512 k-points to 32 atoms with 64 k-points: (64/512) × (32/4)³ = 64 times) | our calculation (the assumptions are stated in the text) | Our calculation |
| Borlido et al.'s figures for all functionals, specific lattice-constant values of Haas et al., the latest progress of the Materials Project recomputation, measured run times of each code, and an international standard for evaluating or reproducing DFT results | not confirmed in the primary sources we could access, so not stated (commentary) | commentary |
| The organisation of the table of what DFT does and does not tell you, how to choose a functional, the table of calculation settings, three habits for reading computed values, how to frame questions for DFT, and designing the experimental confirmation | our own organisation and commentary based on the published content; not views stated by the research groups | commentary |
| Note on figure types: Figs 5, 7, 8 include our calculation, Figs 2, 3, 4, 6 are concept diagrams (vector drawings), and Figs 1, 9 are ImageGen-generated conceptual illustrations | a note by this article (commentary) | commentary |
Last updated: 23 October 2026. Sources are limited to primary material (peer-reviewed papers, arXiv papers and official Materials Project documentation). The numerical examples (atom counts against cost, the supercell example) are our own calculations, not real run times. Computed values are values under stated calculation conditions (functional, settings), not measured values. We could not confirm the figures for all functionals in Borlido et al., the specific lattice-constant values of Haas et al., the latest progress of the Materials Project recomputation, measured run times of each code, or an international standard for evaluating or reproducing DFT, so we do not state them. All figures are concept diagrams or diagrams that include our own calculation; the ImageGen images are conceptual illustrations.