Background
Traditional computational materials science relies heavily on quantum mechanical calculations, which, despite their high accuracy, are computationally expensive and limit the study of large-scale or long-duration phenomena. ML interatomic potentials offer a compelling solution by providing near-first-principles accuracy at a fraction of the computational cost. This enables researchers to explore complex material behaviors, analyze defects, and predict material durability across various applications, from energy storage to catalysis and semiconductors.
Key Findings
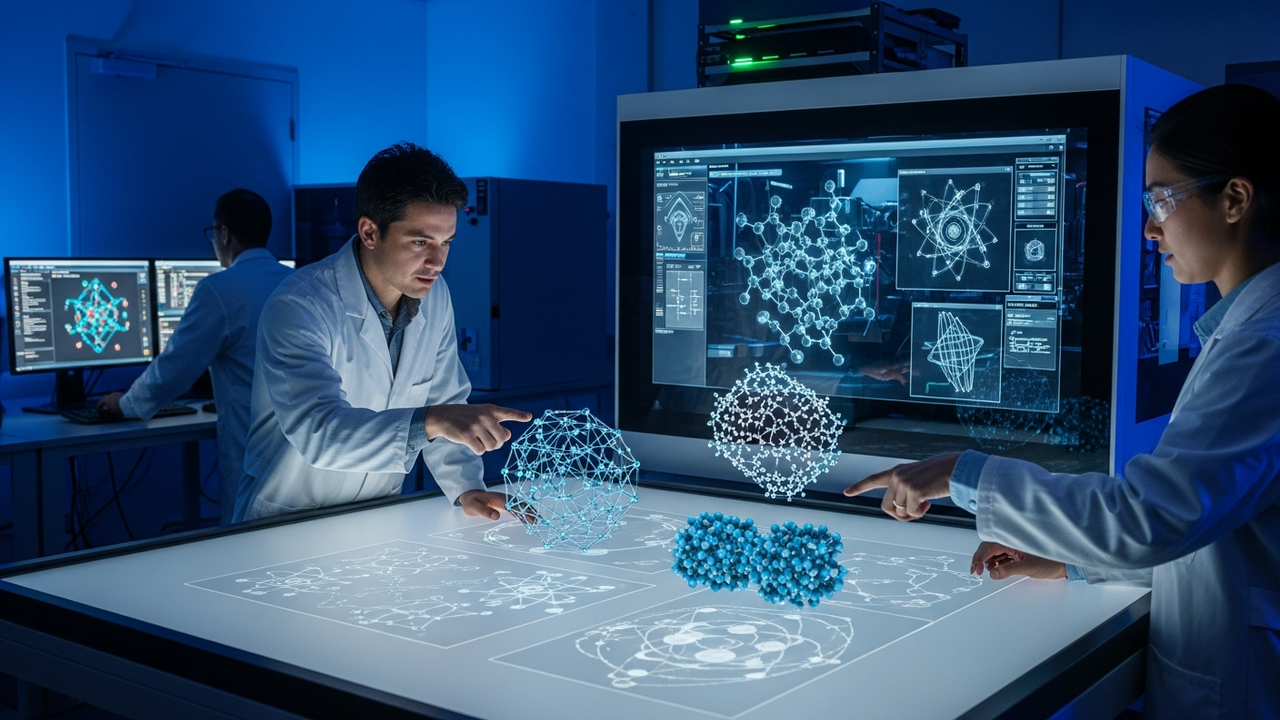
The AI4Science Seminar at Chalmers University of Technology highlighted the transformative role of machine learning (ML) in computational chemistry and materials research, specifically emphasizing molecular characterization and ML interatomic potentials. The discussions underscored the critical need for automated training pipelines for ML interatomic potentials and the creation of large, high-quality materials databases, positioning them as foundational elements for next-generation materials science.
The seminar further elaborated on significant advancements and future directions in applying ML to fundamental scientific problems. A core theme was the development and application of ML interatomic potentials, designed to mimic the accuracy of quantum chemical calculations (first-principles methods) with significantly reduced computational cost, thus allowing for the simulation of larger systems and longer timescales than previously possible.
- Automated Training Pipelines (e.g., autoplex): Speakers emphasized the development of automated pipelines for training ML interatomic potentials. These systems can autonomously generate and refine potentials from first-principles data, dramatically reducing the manual effort and expertise required. This automation accelerates the iteration cycle of potential development, leading to more robust and accurate models.
- Large-Scale, High-Quality Material Databases: The performance of ML models is intrinsically linked to the quality and quantity of their training data. The seminar highlighted the importance of curating and expanding large, high-quality materials databases, potentially augmented by generative models. These databases serve as the bedrock for training advanced ML models, ensuring their reliability and generalizability across diverse material systems.
- Enhanced Molecular Characterization: ML models are increasingly being used for rapid and accurate prediction of molecular properties, including electronic structure, thermodynamic behavior, and reaction pathways. This capability significantly streamlines the screening process for novel materials and drug candidates.
The event convened experts from quantum chemistry, machine learning, and condensed matter physics, fostering interdisciplinary discussions on how these cutting-edge technologies can collectively accelerate scientific discovery.
The push for automated training of ML interatomic potentials and the expansion of high-quality materials databases are set to redefine the landscape of computational materials science. This will empower researchers to simulate and predict the behavior of complex material systems at unprecedented scales. In the near future, ML-driven simulations are expected to become a primary engine for new material discovery, playing a crucial role in accelerating product development cycles across industries such as energy storage, catalysis, semiconductor manufacturing, and biomaterials. This strategic integration of AI holds the potential to unlock breakthroughs that were previously computationally intractable.
Source: https://psolsson.github.io/AI4ScienceSeminar
Get our weekly technology intelligence — free
Receive an infographic that lets you judge at a glance whether each field’s analysis report is worth reading.
Subscribe Free — Weekly Tech Intelligence
By subscribing, you’ll receive Troy-Technical’s weekly technology intelligence newsletter.
- Your email and selected fields are used only to deliver the newsletter.
- We never share your information with third parties.
- You can unsubscribe anytime via the link in each email.
See our Privacy Policy for details.
Takes about a minute · Unsubscribe anytime

Comments